Evolution and conservation genomics
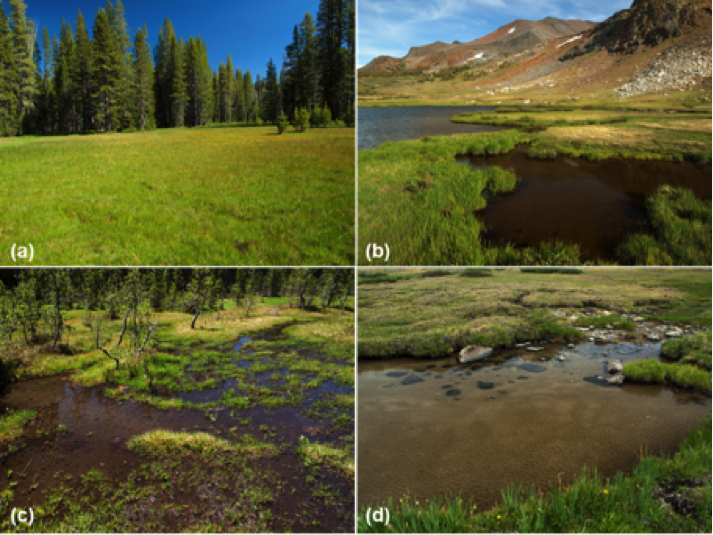
Almost every Yosemite toad breeds in a wet meadow, in pools that appear when the snow melts and are gone by late summer. The meadows are discrete and mappable, separated by dry forest and granite, so the units of population structure can be pointed at rather than inferred. Pleistocene glaciers split and rejoined lineages here repeatedly, leaving replicate contact zones within a few kilometres of each other. The species was also among the first amphibians whose collapse was documented in detail. Divergence, admixture, and adaptation are all legible within one small range, in a federally threatened animal whose habitat is moving upslope.
How an ancient process could explain the present
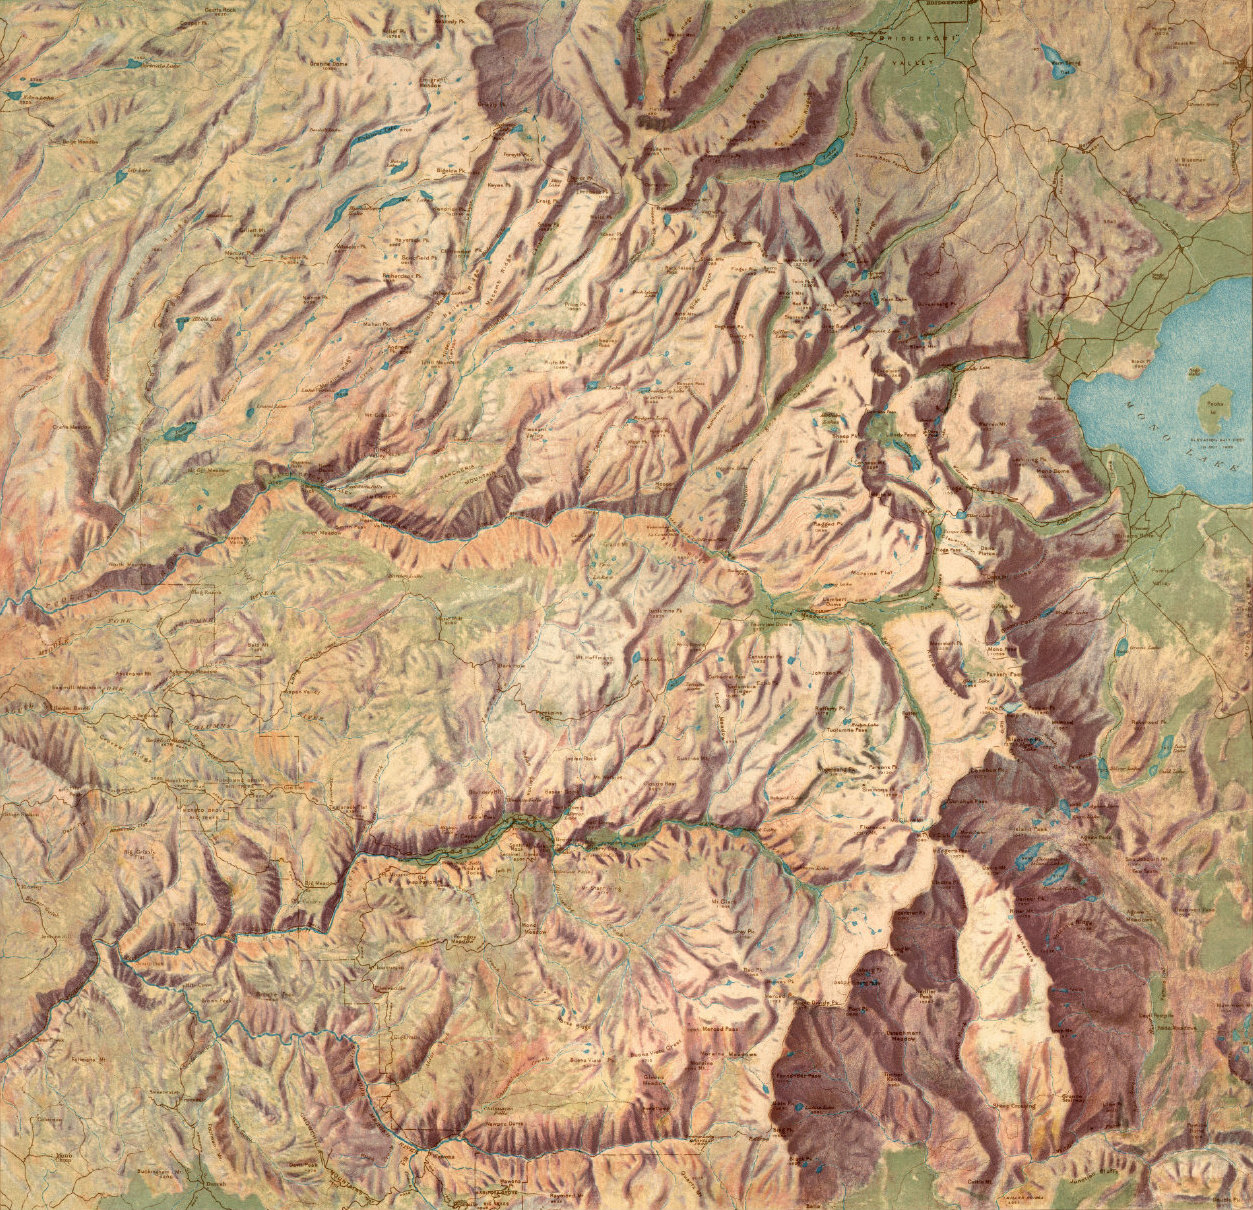
Near the end of the Pliocene, the Sierra Nevada underwent some dramatic changes. Tectonic action that was already well underway began accelerating in earnest, raising the mountains to new heights. By the time the Pleistocene began around two million years ago, the peaks loomed imposingly over the San Joaquin Valley. Glaciers began to advance and recede in roughly 41,000-year cycles, isolating animals and plants into refugia. More recently that periodicity shifted to 100,000-year cycles, when the eccentricity of Earth’s orbit changed.
The biogeographic consequences were enormous. Over 73% of amphibian and squamate species that transect the Sierra Nevada show lineage divergence in this ecoregion.
Species endemic to alpine environments often evolve along steep ecological selection gradients between lowland and upland habitat. But alpine environments also faced repeated glaciation, fracturing those endemics into isolated populations. I introduced a glacial pulse model of alpine speciation, in which cycles of allopatry and ecologically divergent glacial refugia both play a role in generating biodiversity.
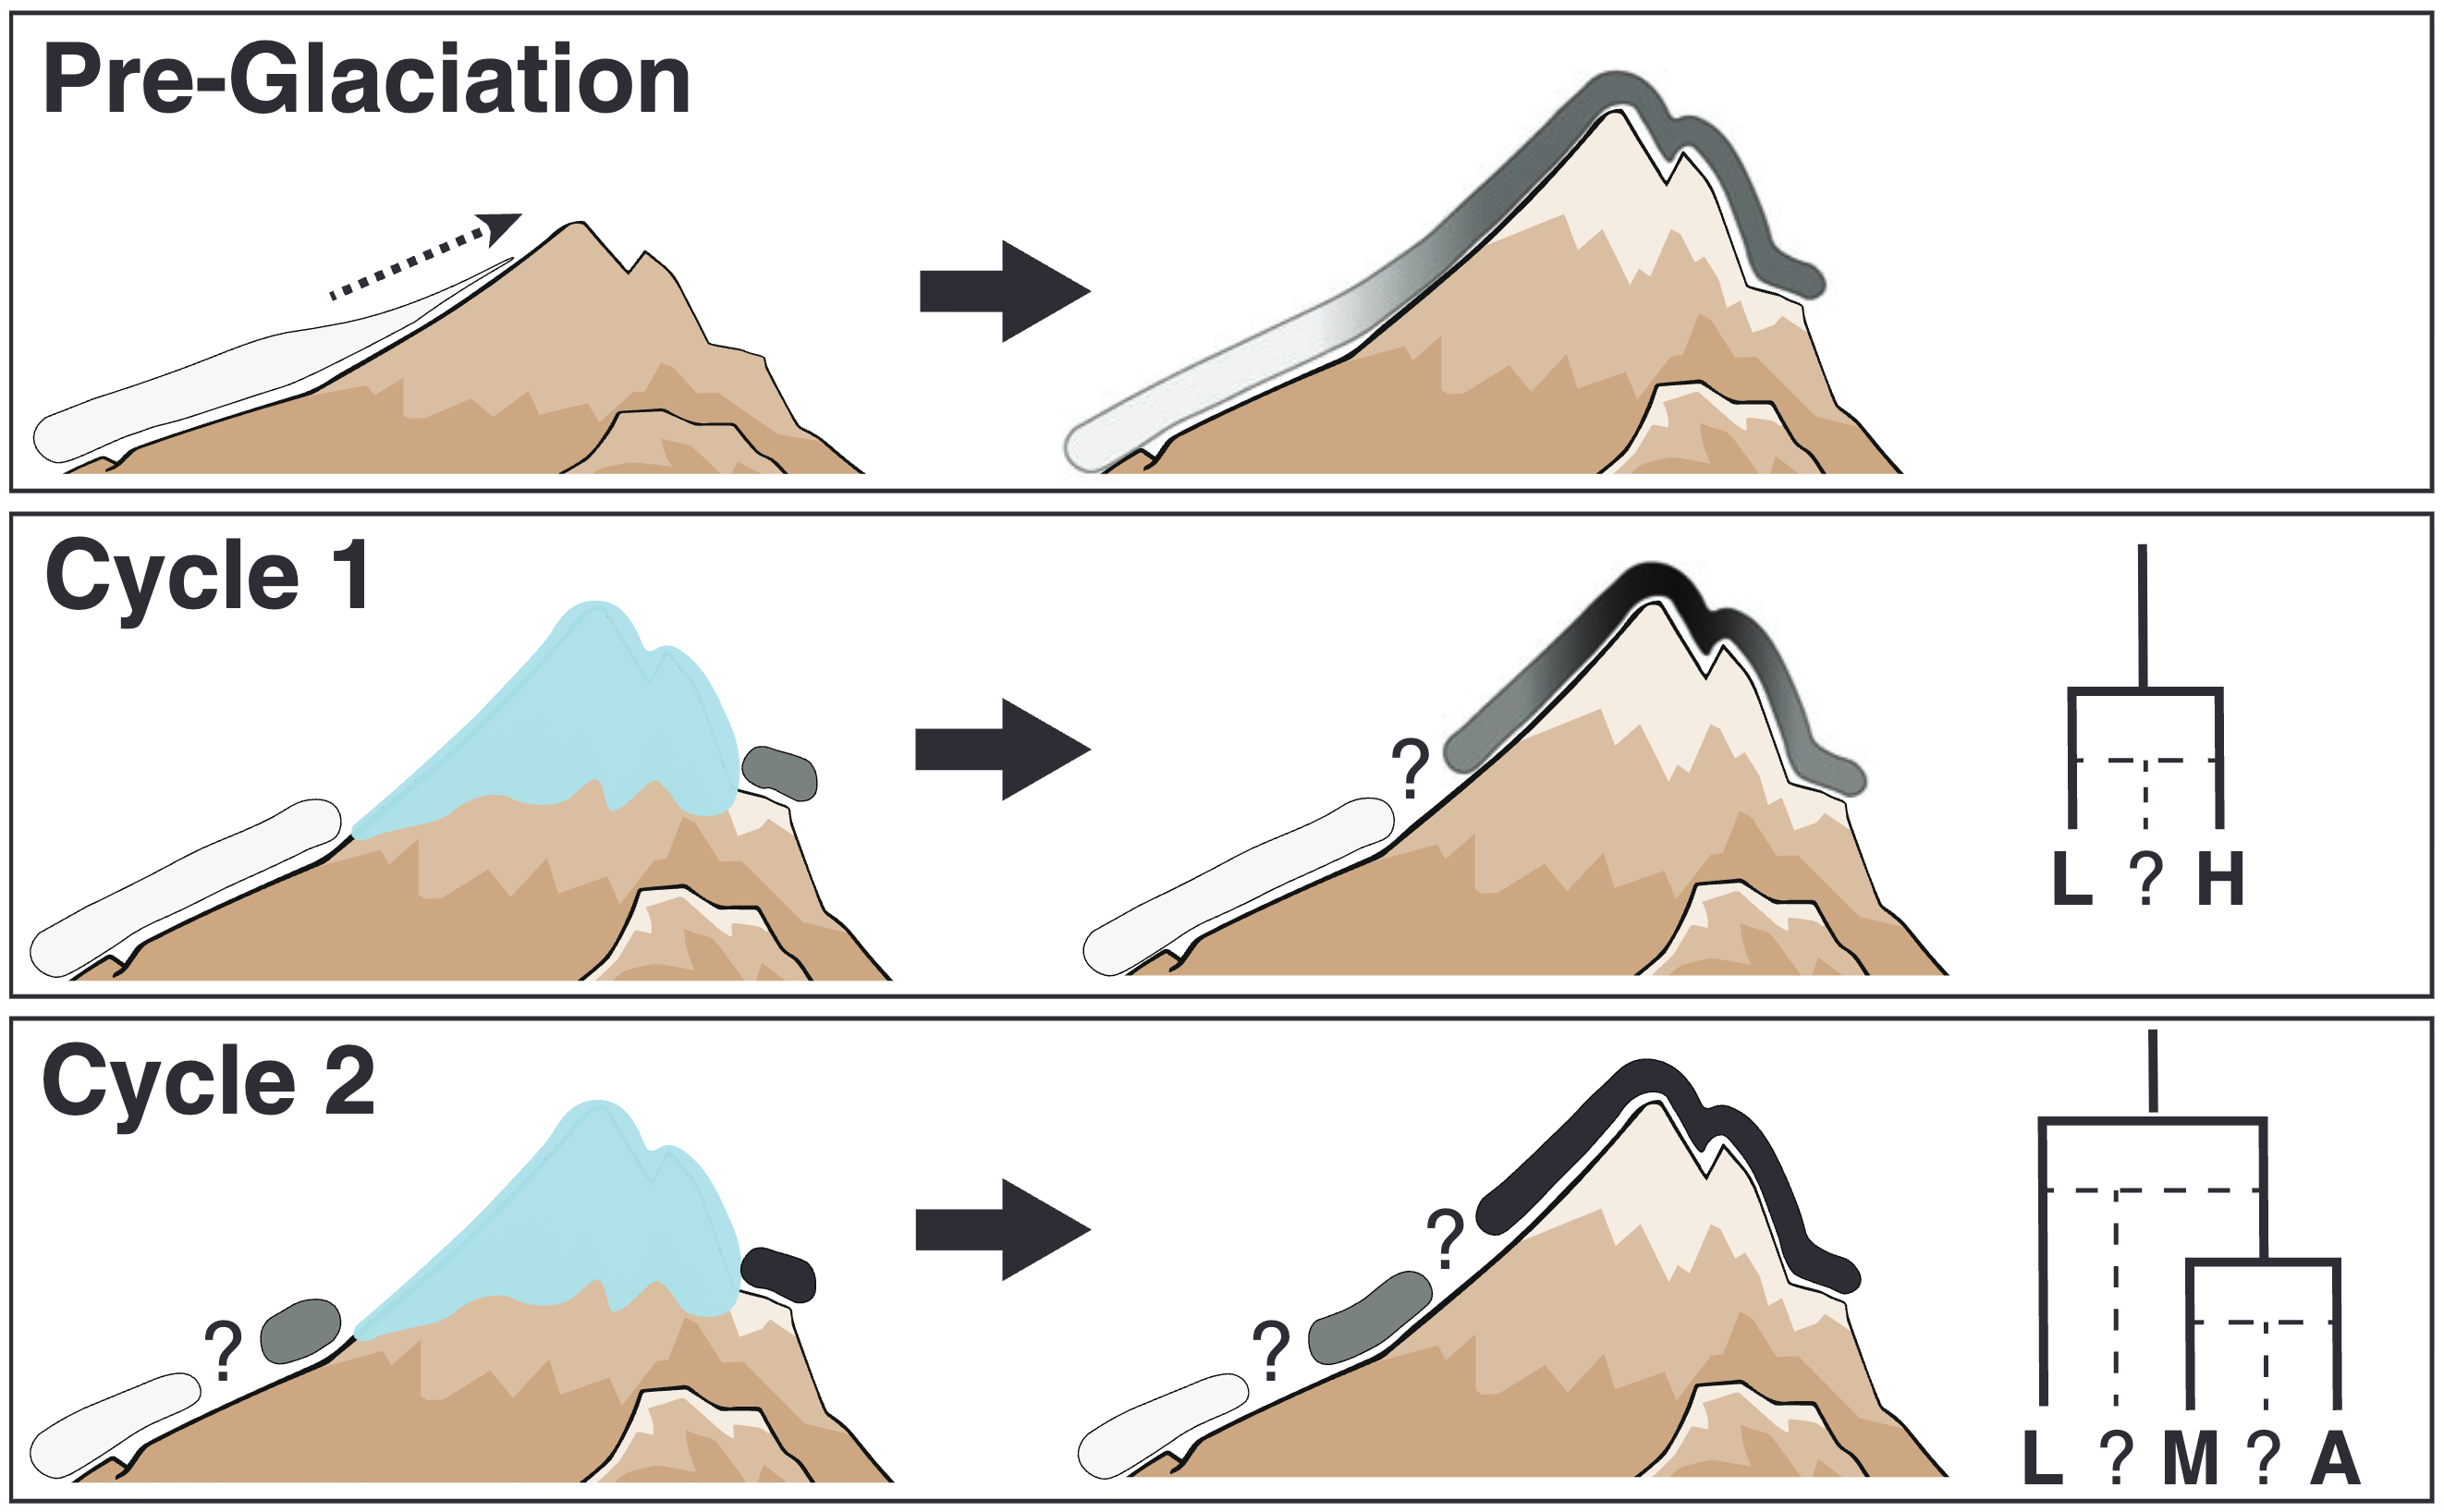
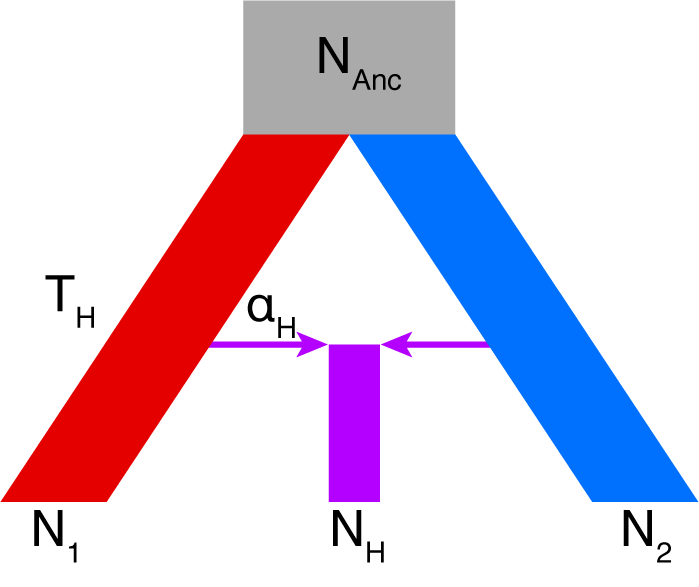
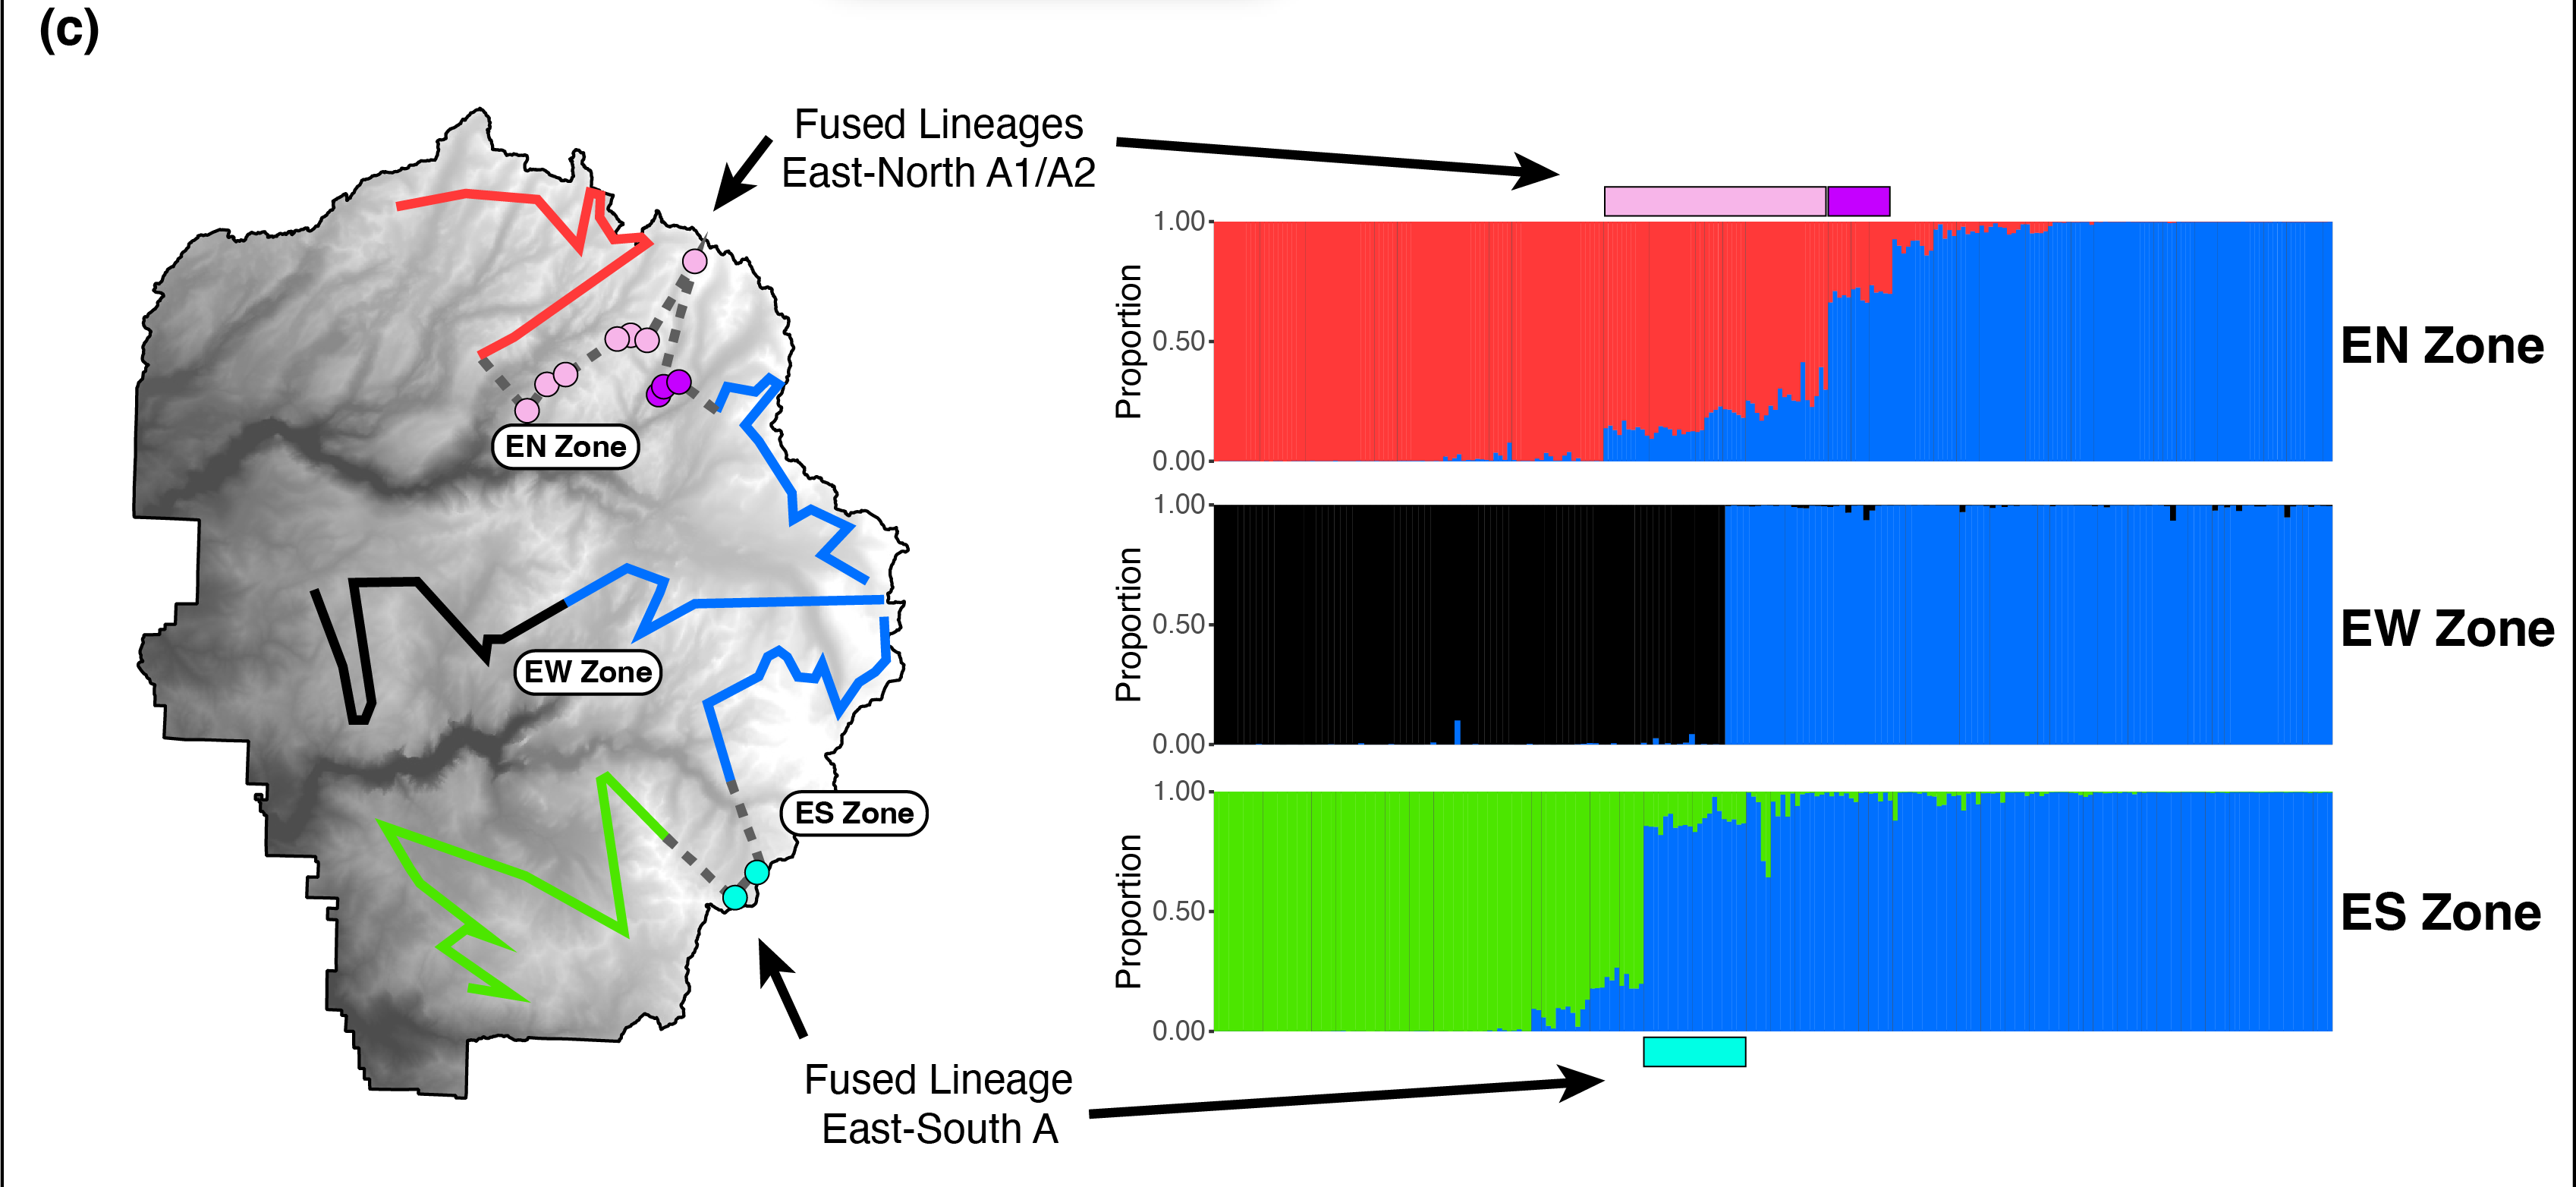
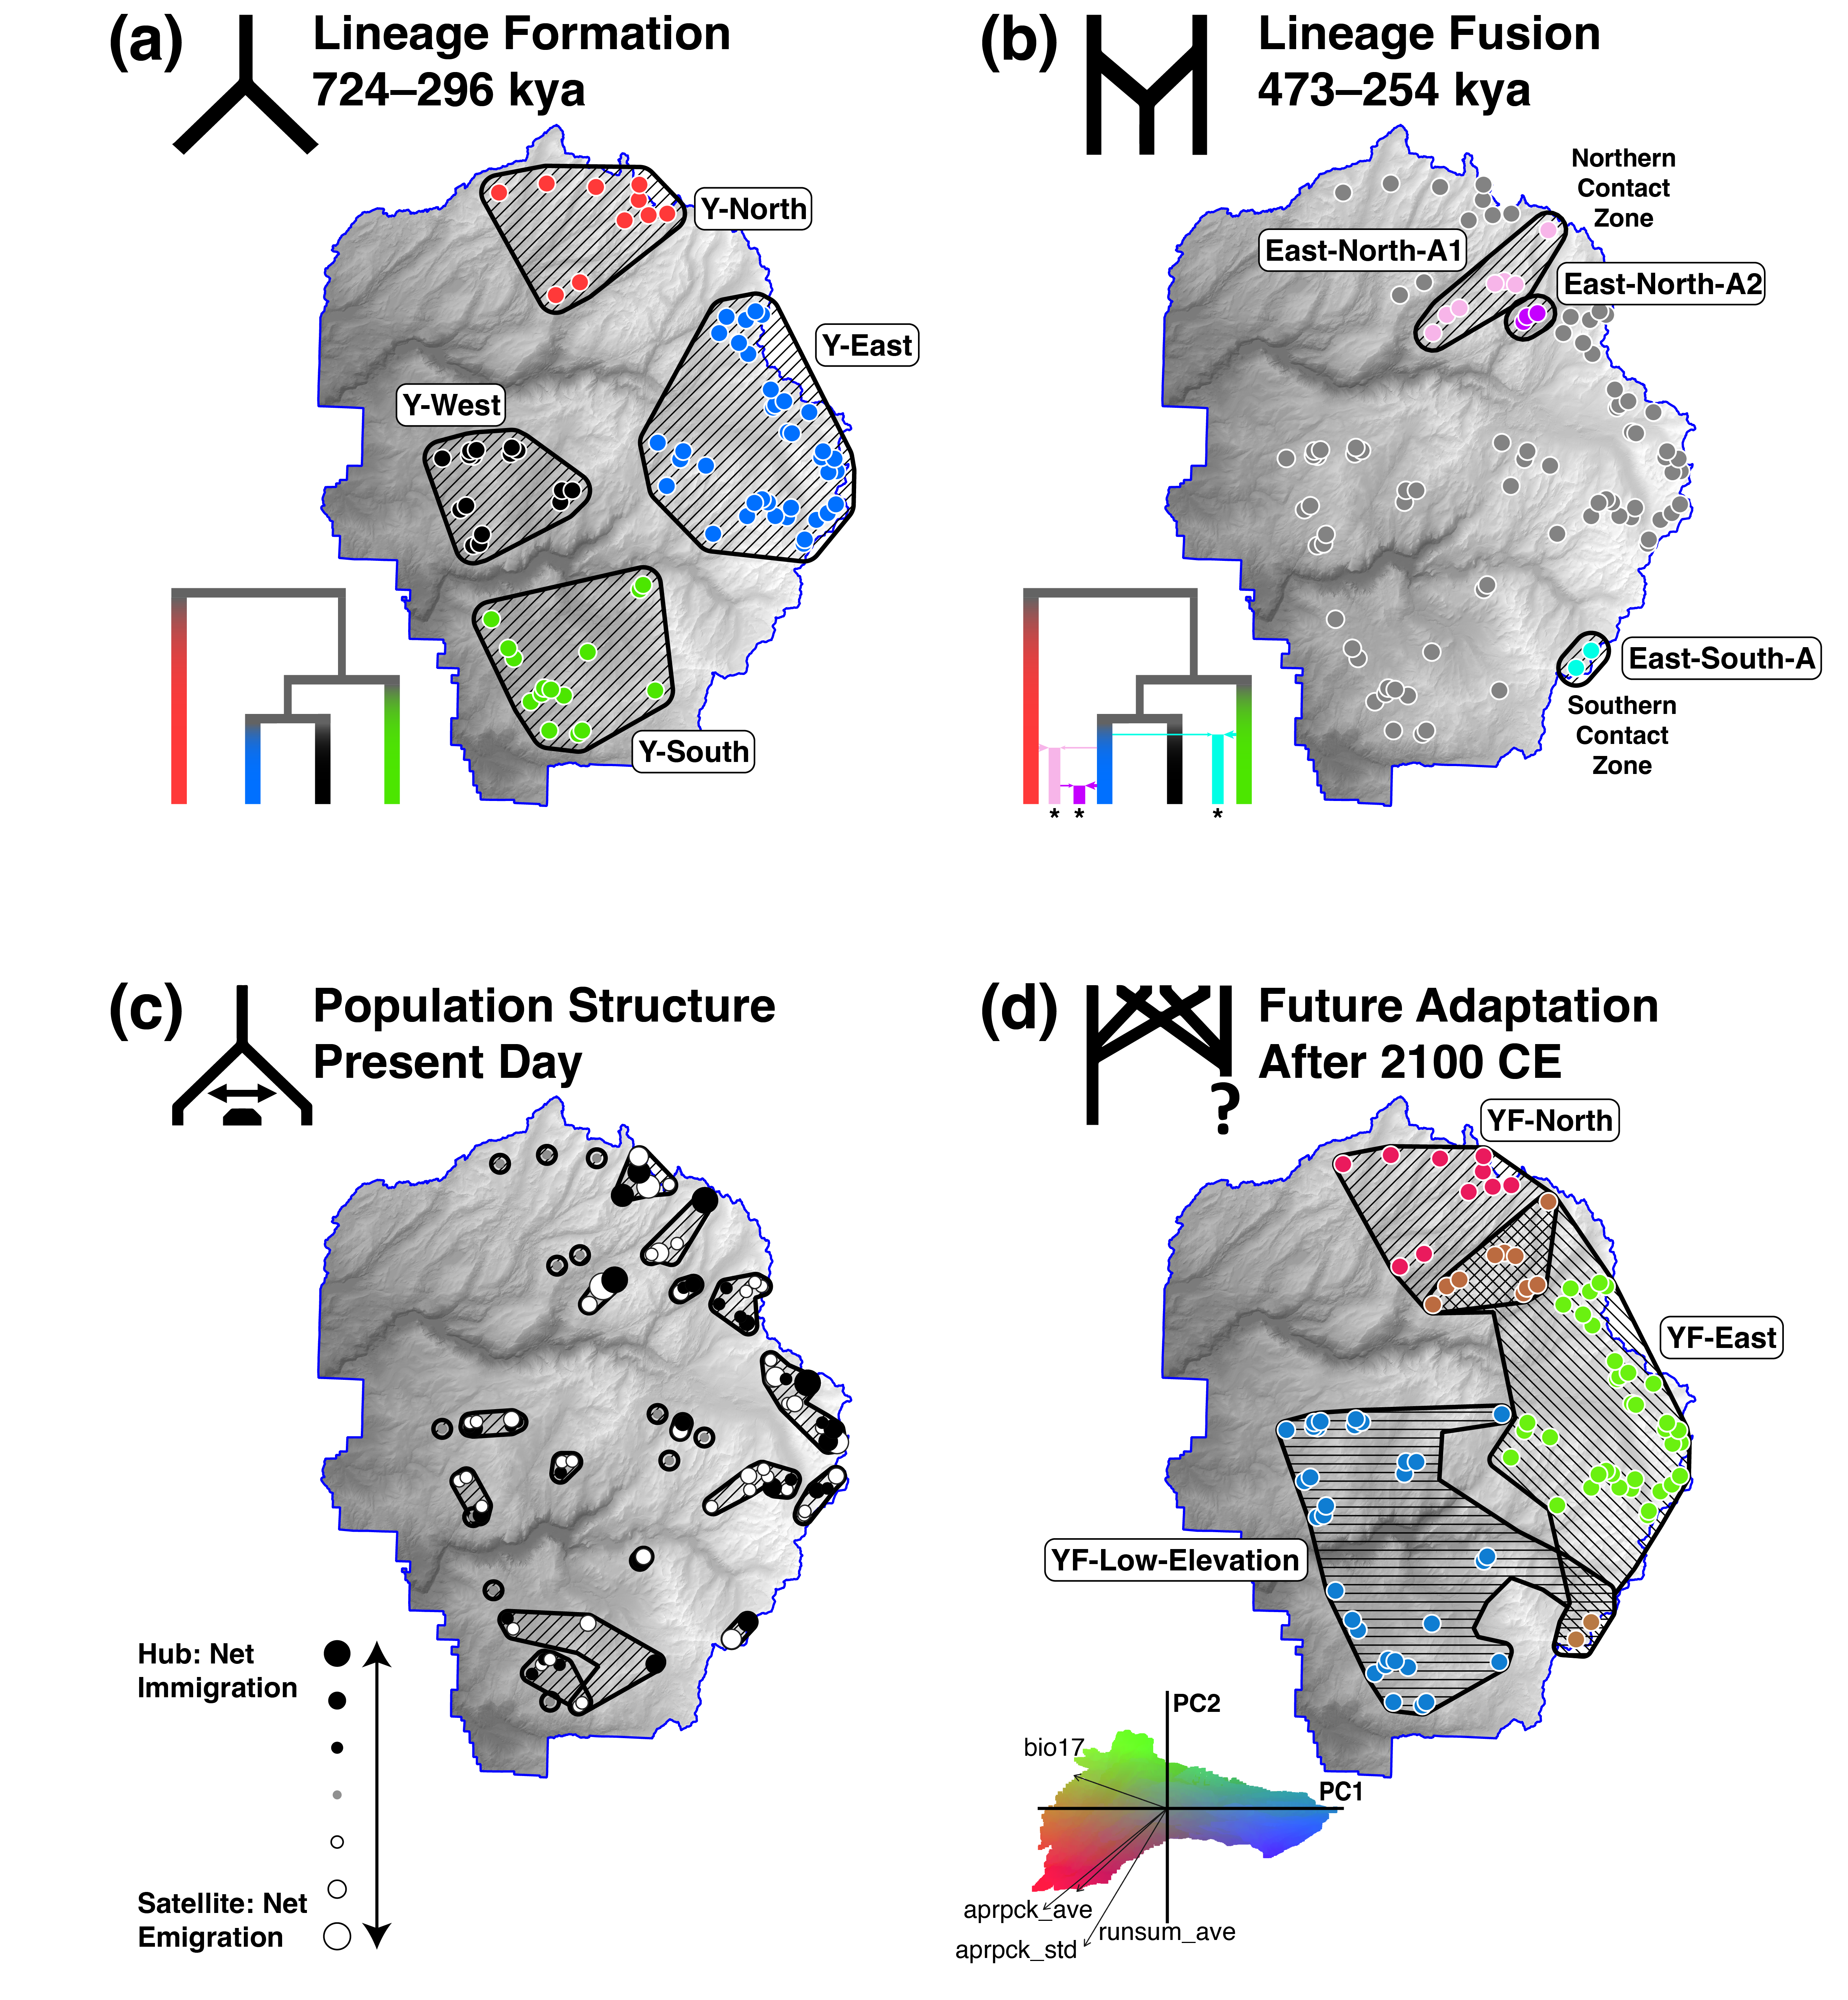
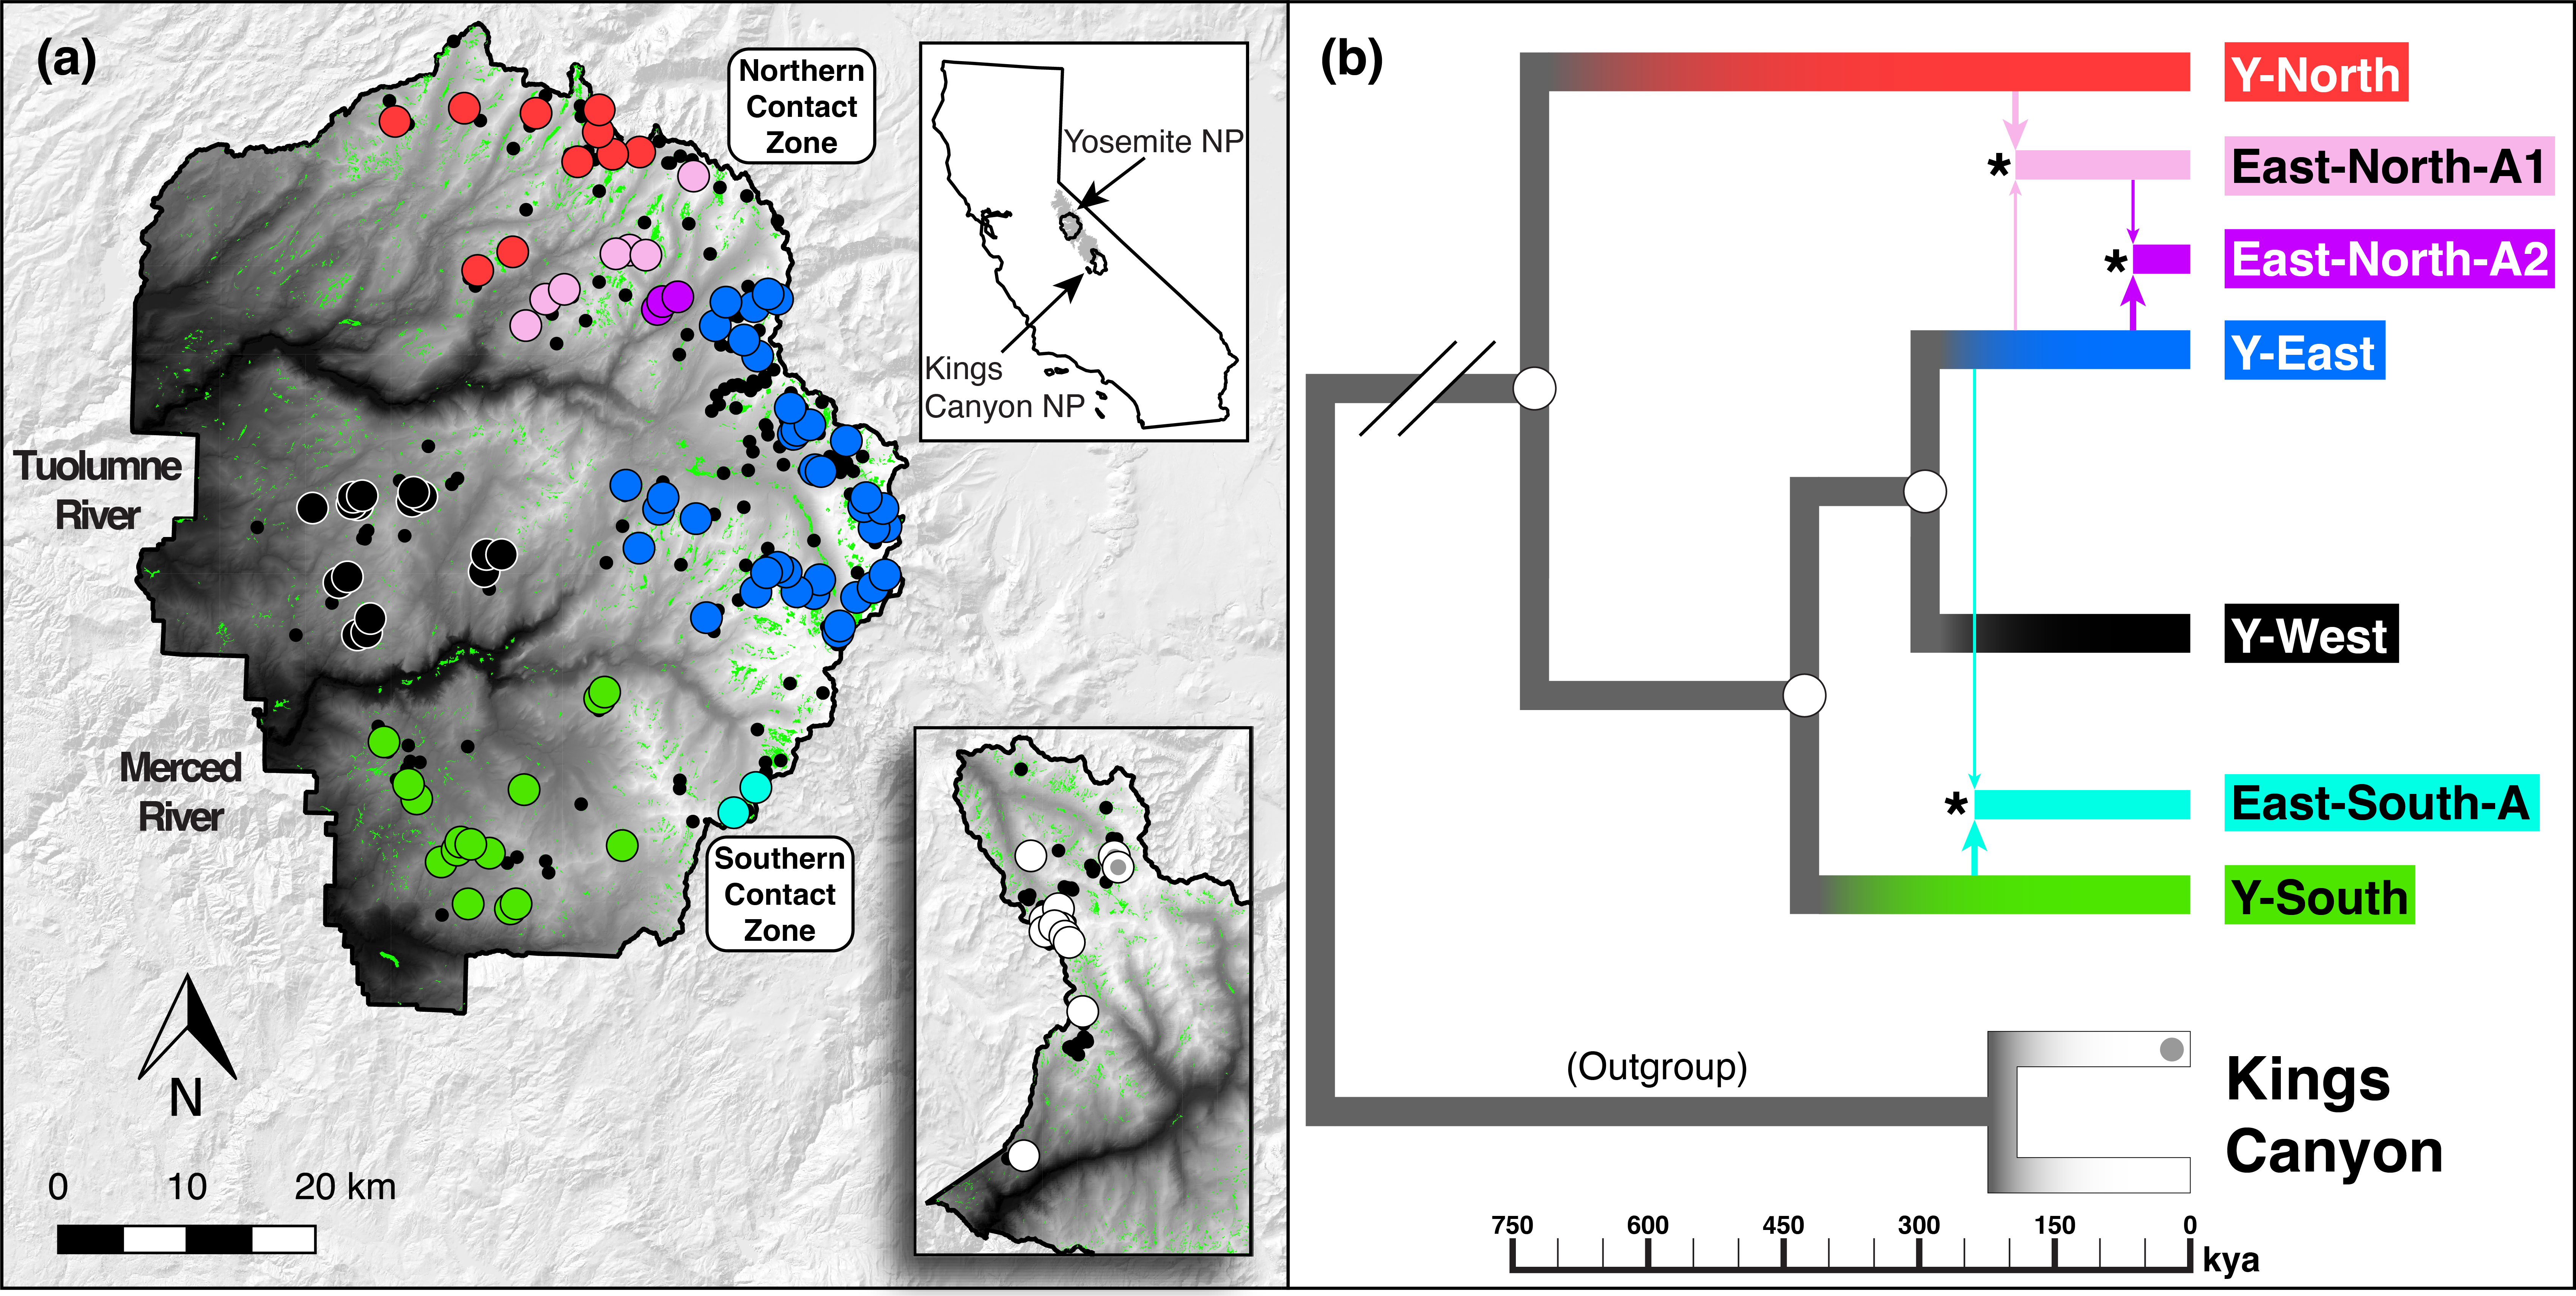
Using double-digest RADseq across a major portion of the species range, we identified nine distinct lineages with divergence times ranging from 18 to 724 thousand years ago, coinciding with multiple Sierra Nevada glacial events. Three of those lineages are products of lineage fusion, meaning two other lineages produced them by admixture. Demographic models suggest the fused lineages have persisted through past glacial cycles, so they are viable despite their hybrid origin.
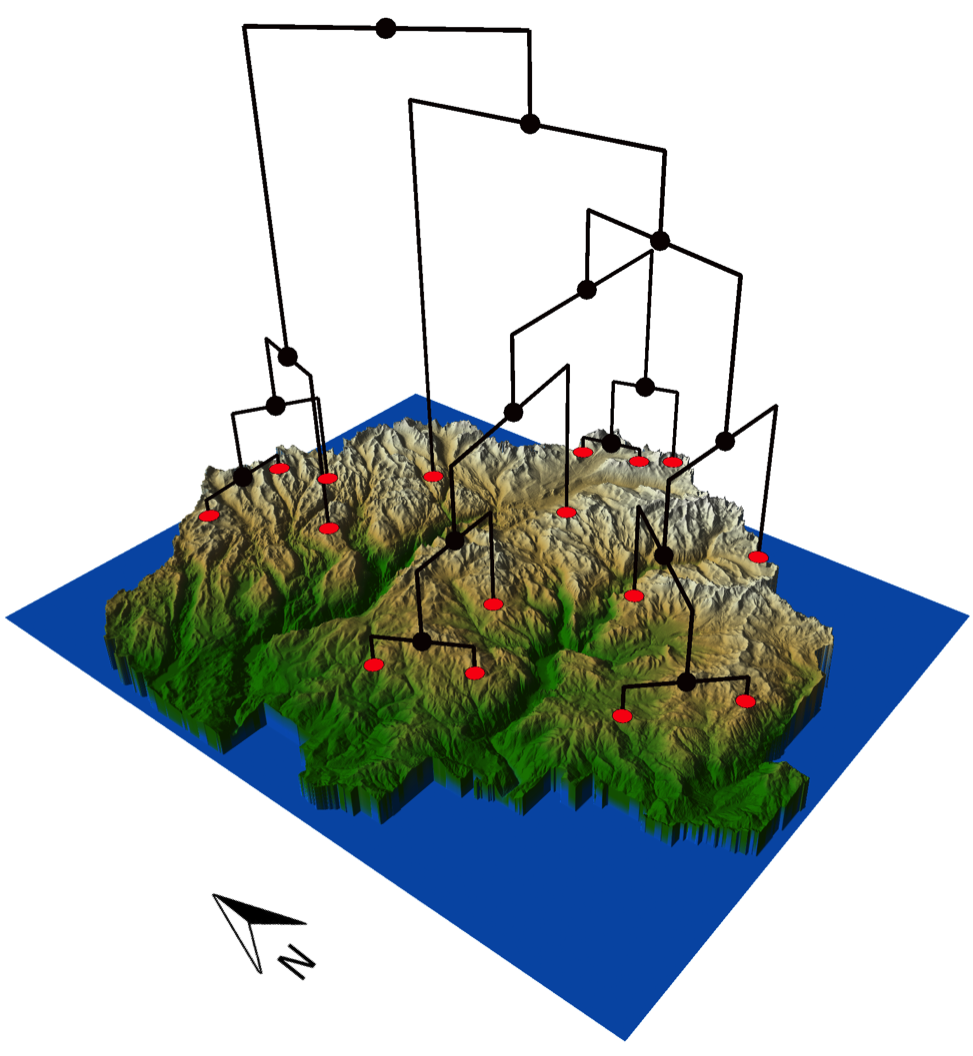
Multiple measures of genealogy shape supported the hypothesis that some lineages recolonized Yosemite from east of the ice sheet, while others remained in western refugia. We also found evidence that low- and high-elevation lineages have repeatedly adapted to divergent climatic niches. Several pairs of lineages were isolated into low western and high eastern refugia, and they do not appear to interbreed freely on secondary contact.
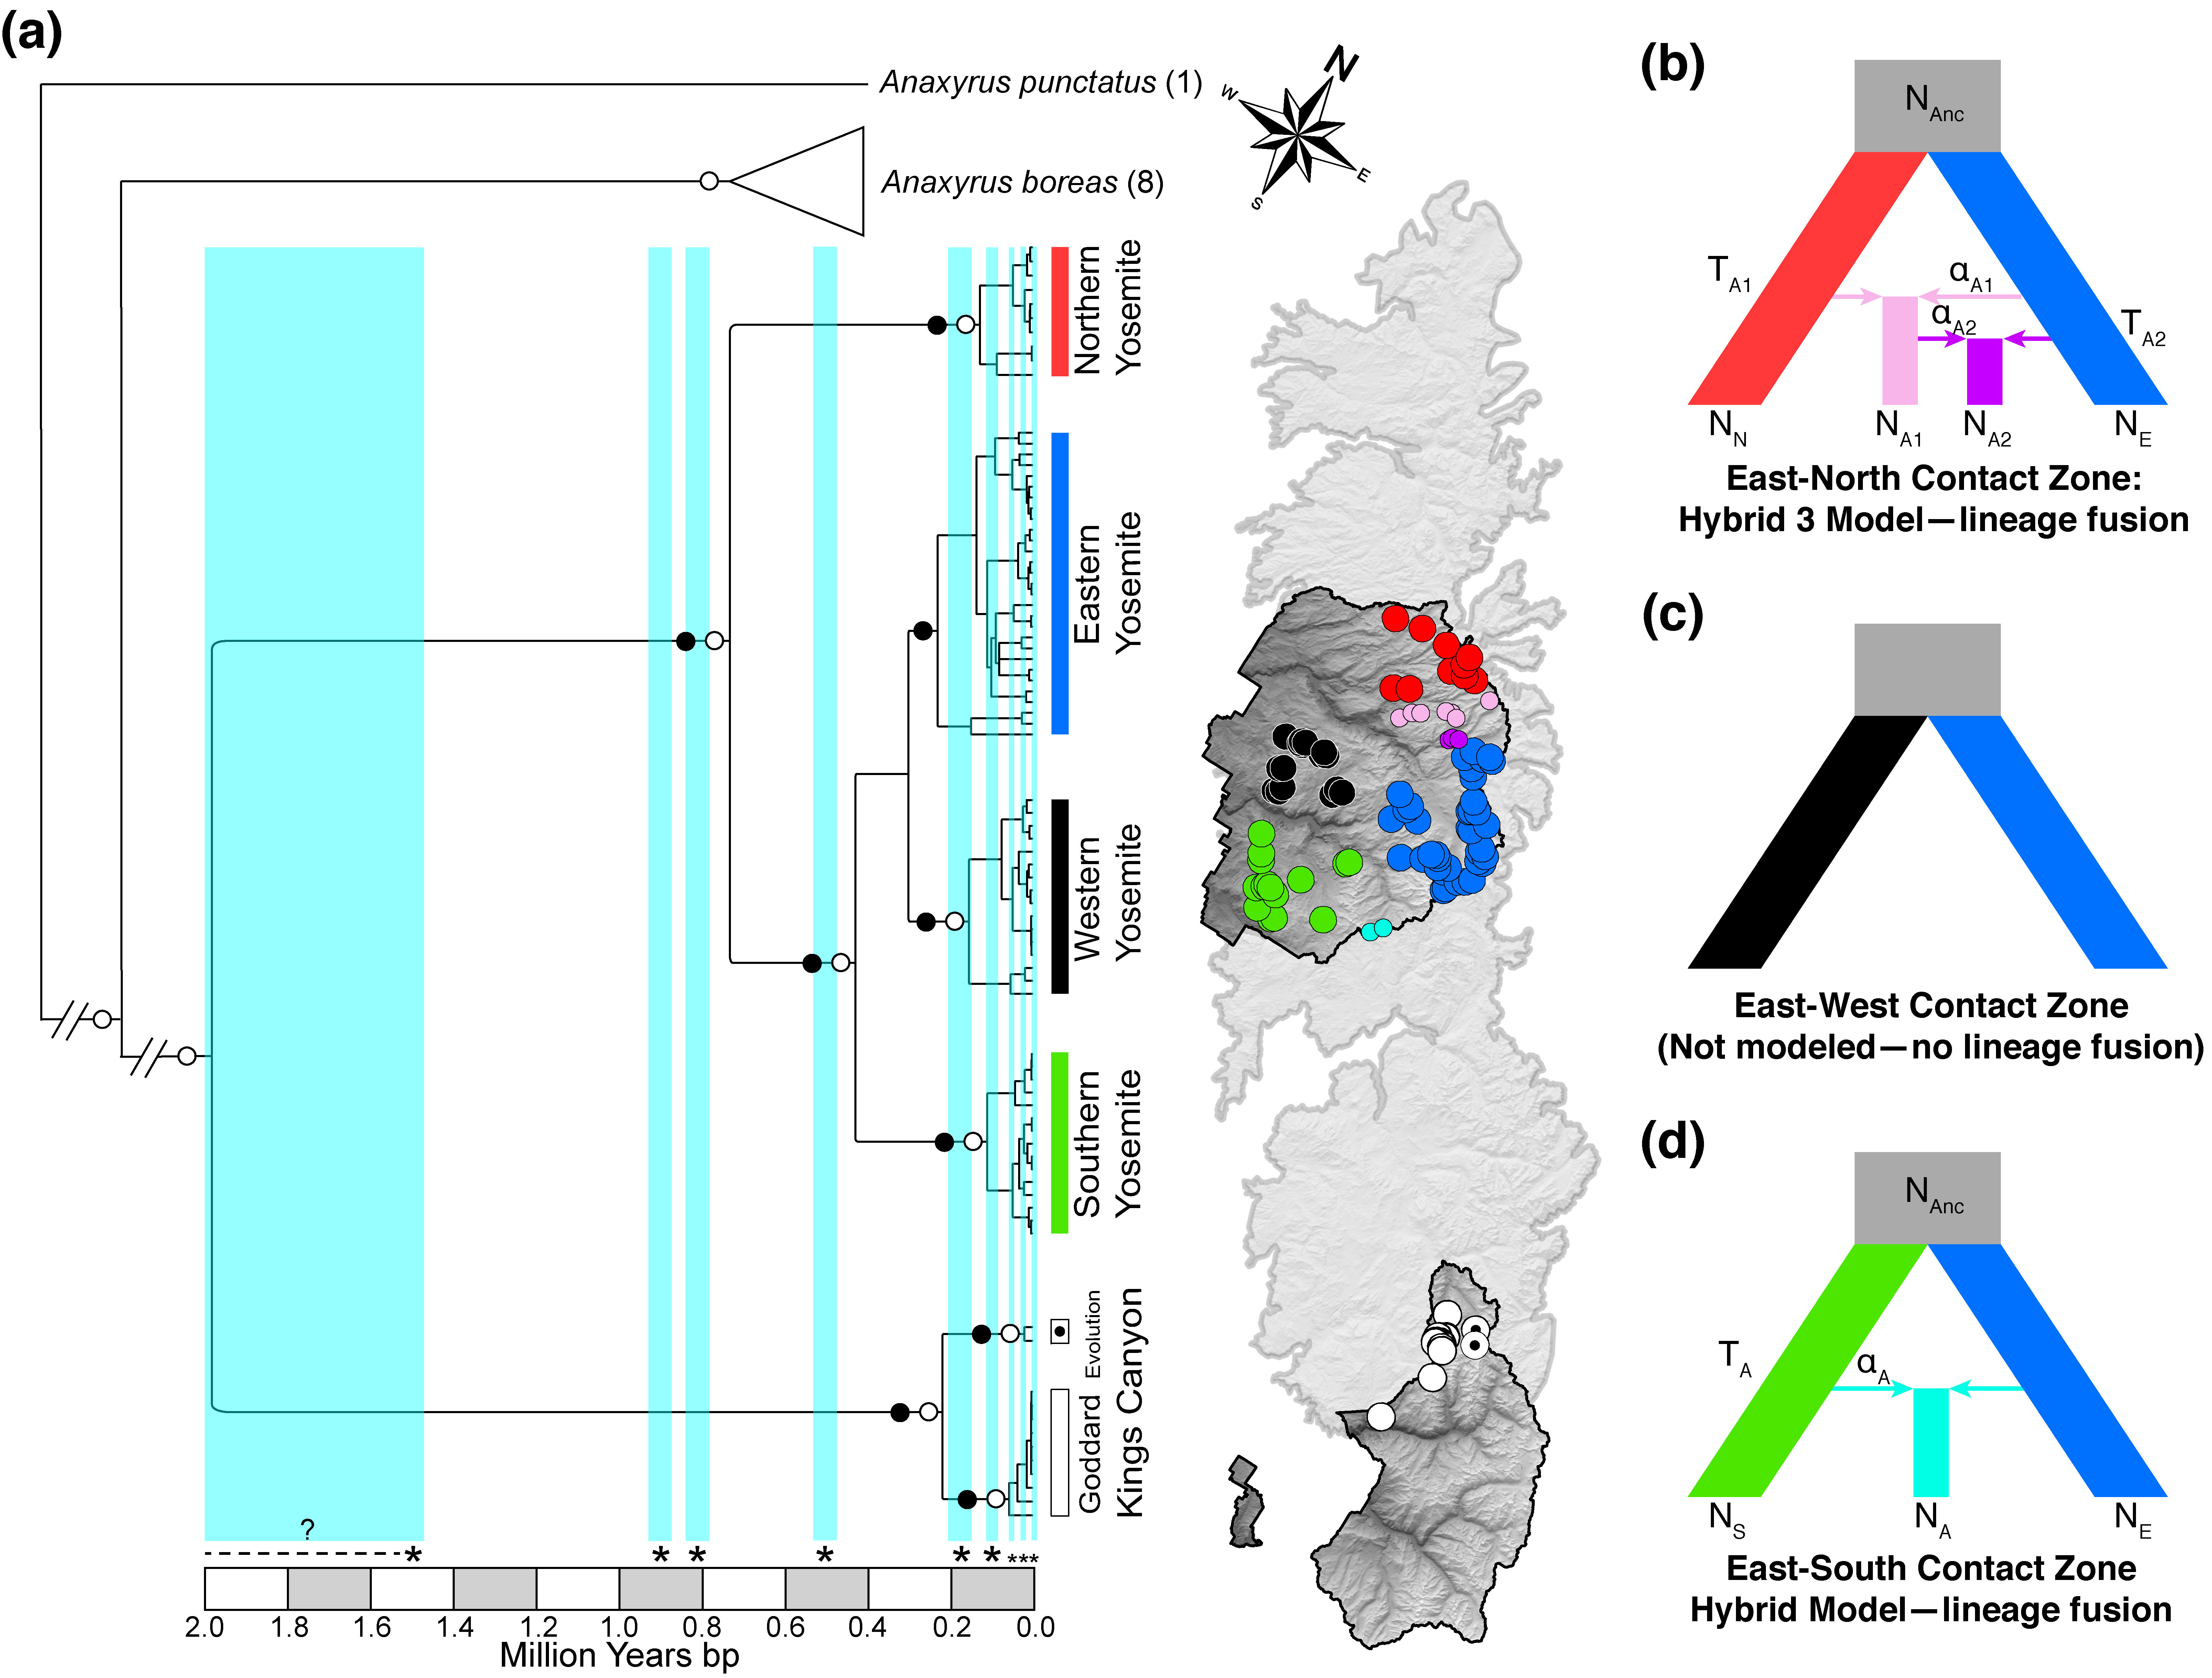
1. Lineage divergence has involved both allopatry and ecological divergence, not one or the other.
2. Lineage fusion via secondary contact is an under-appreciated phenomenon. Fused lineages could be important crucibles of adaptive diversity across deep evolutionary time.
Technical details
This project was a large undertaking. Using double-digest RAD sequencing I genotyped 650+ individuals at 3,000+ RAD haplotype loci. Seven lanes of an Illumina HiSeq 2500 returned 1.88 billion reads, comprising 161.68 Gb of usable sequence at a mean quality score of 35.57. That is over fifty times the size of the human genome.
Traditional pipelines pick one SNP per RAD locus and discard the rest, which throws away a great
deal of information. I wrote an extensive Python tool,
fasta2genotype.py, that uses
haplotype information from entire sequences instead.
Analyses included concatenated and species-coalescent phylogenetics, species network analysis, demographic modeling from the joint site frequency spectrum, spatial modeling from the SFS, Bayesian skyline analysis, MaxEnt reconstruction of glacial refugia, and tests of niche divergence. Divergence times came from a BEAST chronogram calibrated on a molecular rate.
Where the gene pools actually begin and end
Meadow neighborhoods, hubs, and satellites
Before you can ask how climate will affect connectivity, you need to know what the units of connectivity are. Toads move between meadows, but not evenly and not symmetrically, and the boundaries of a gene pool are not obvious from a map.
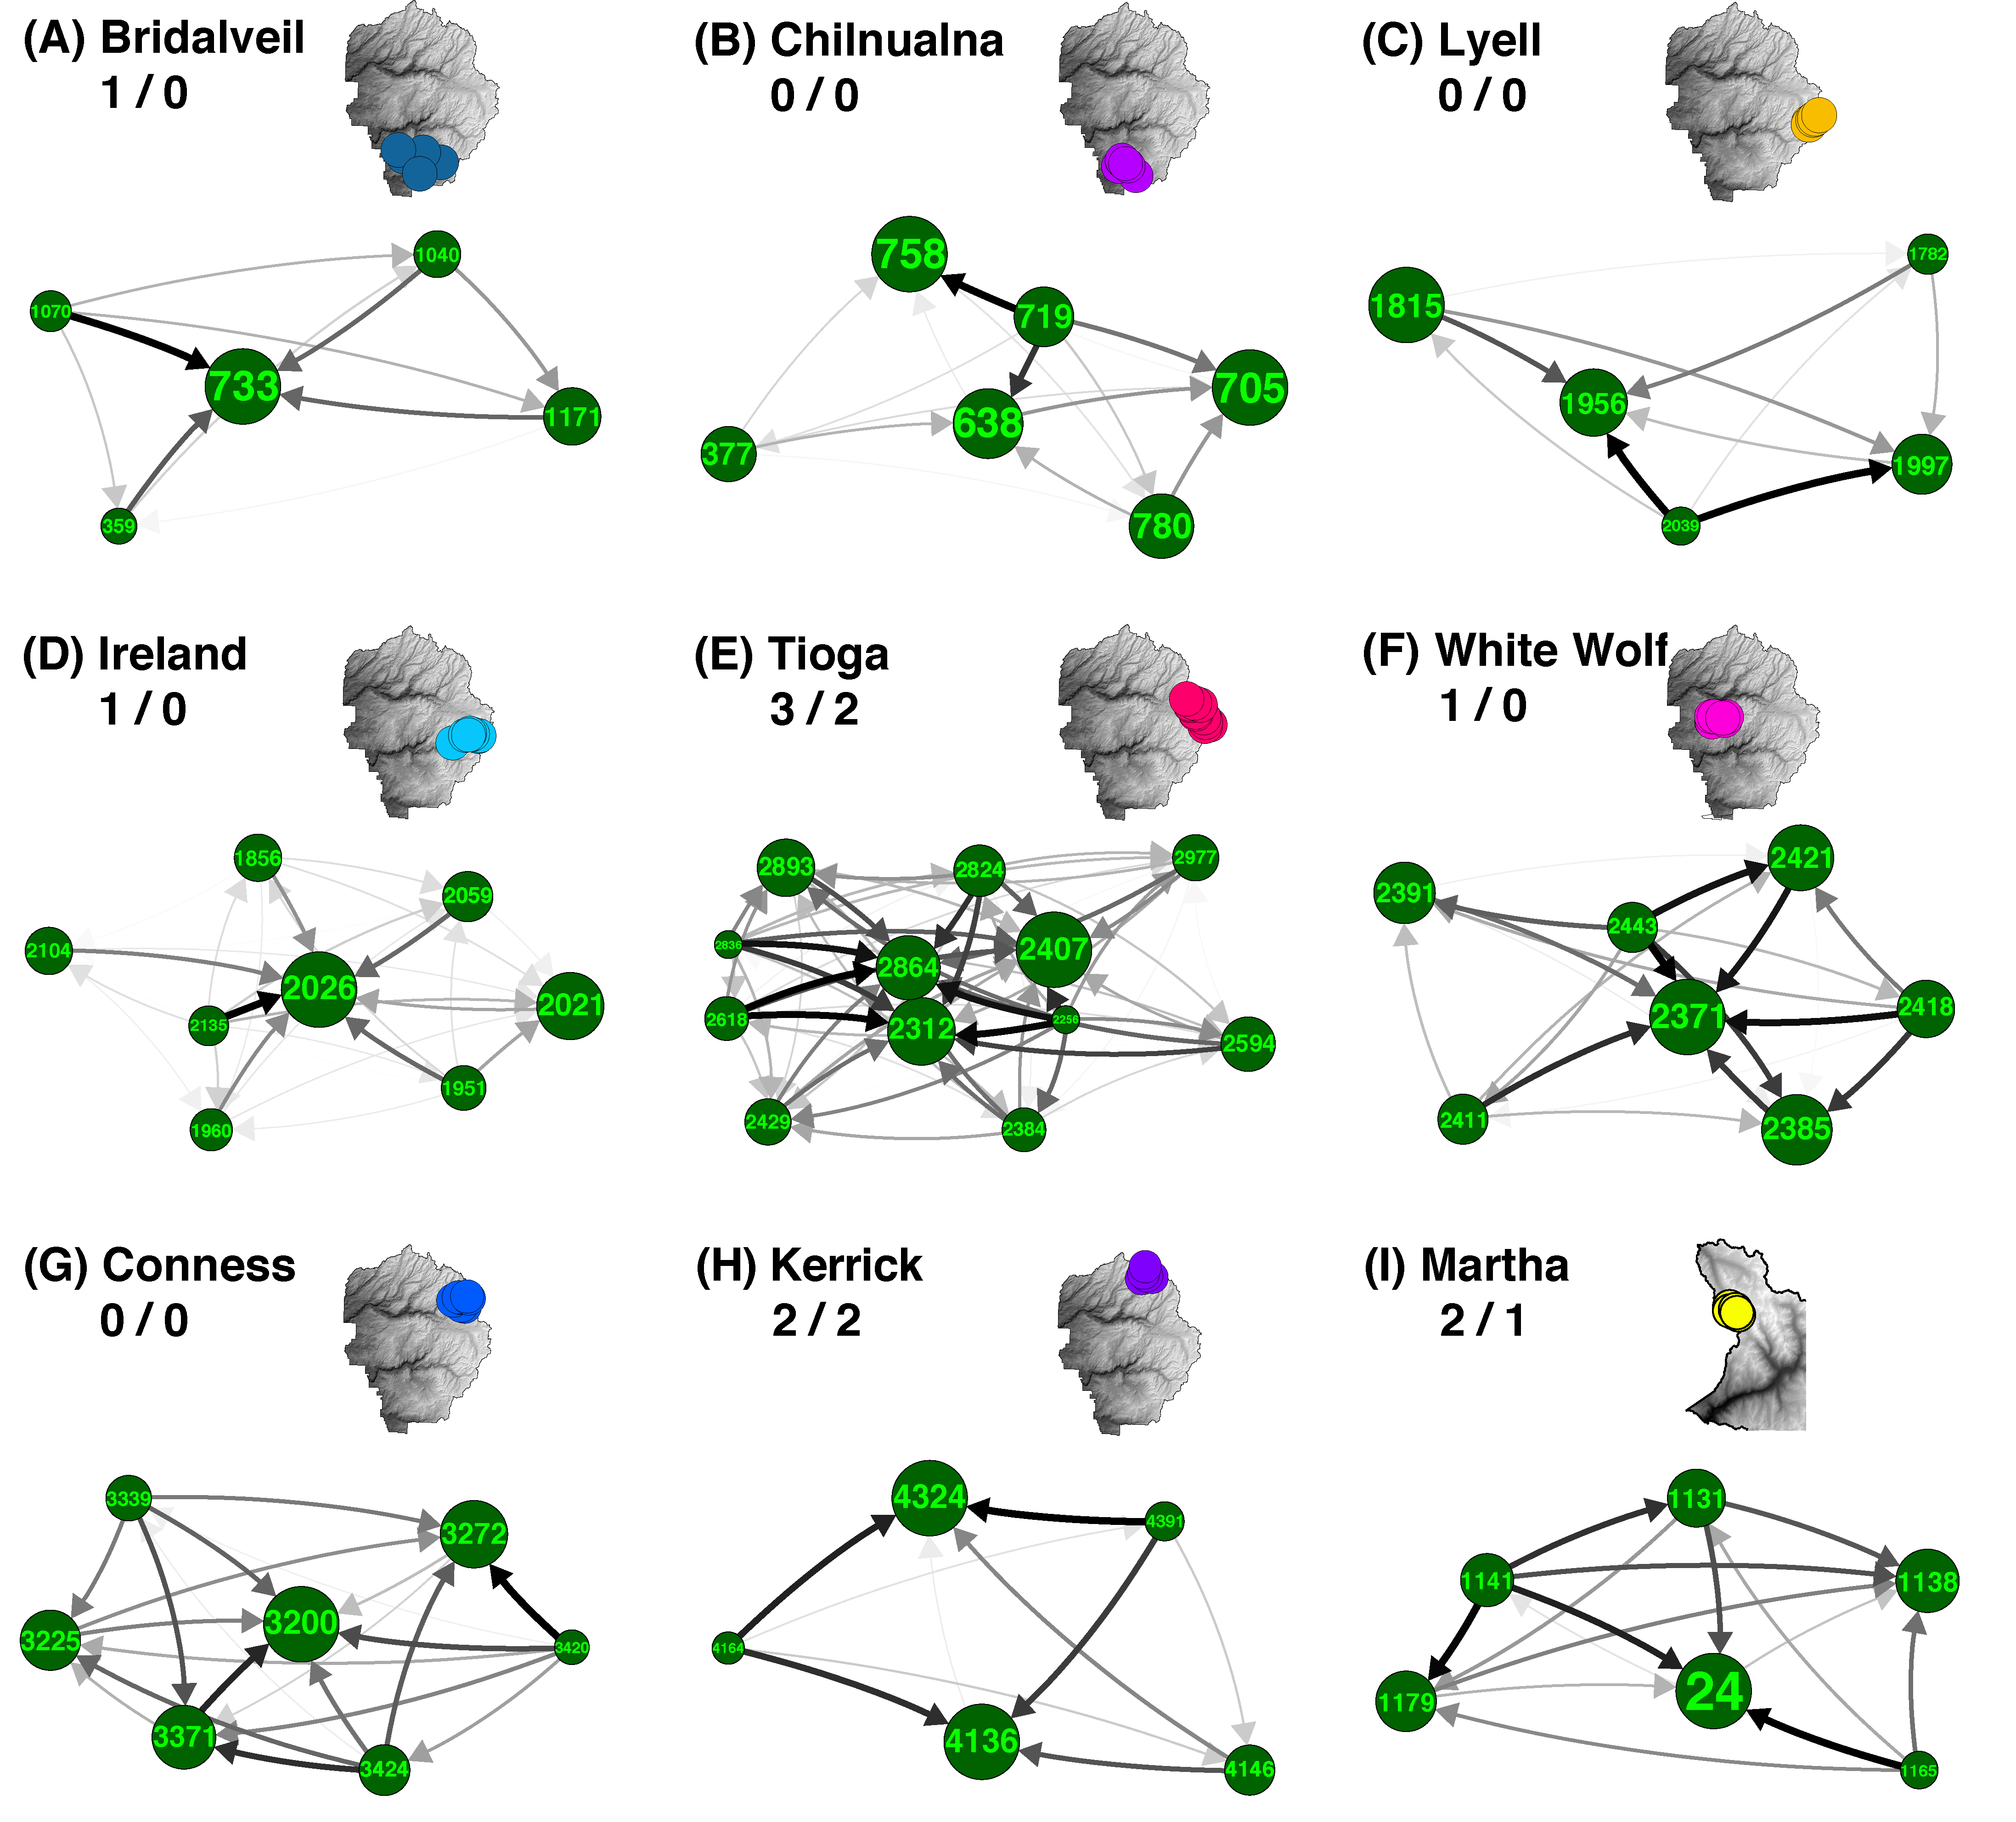
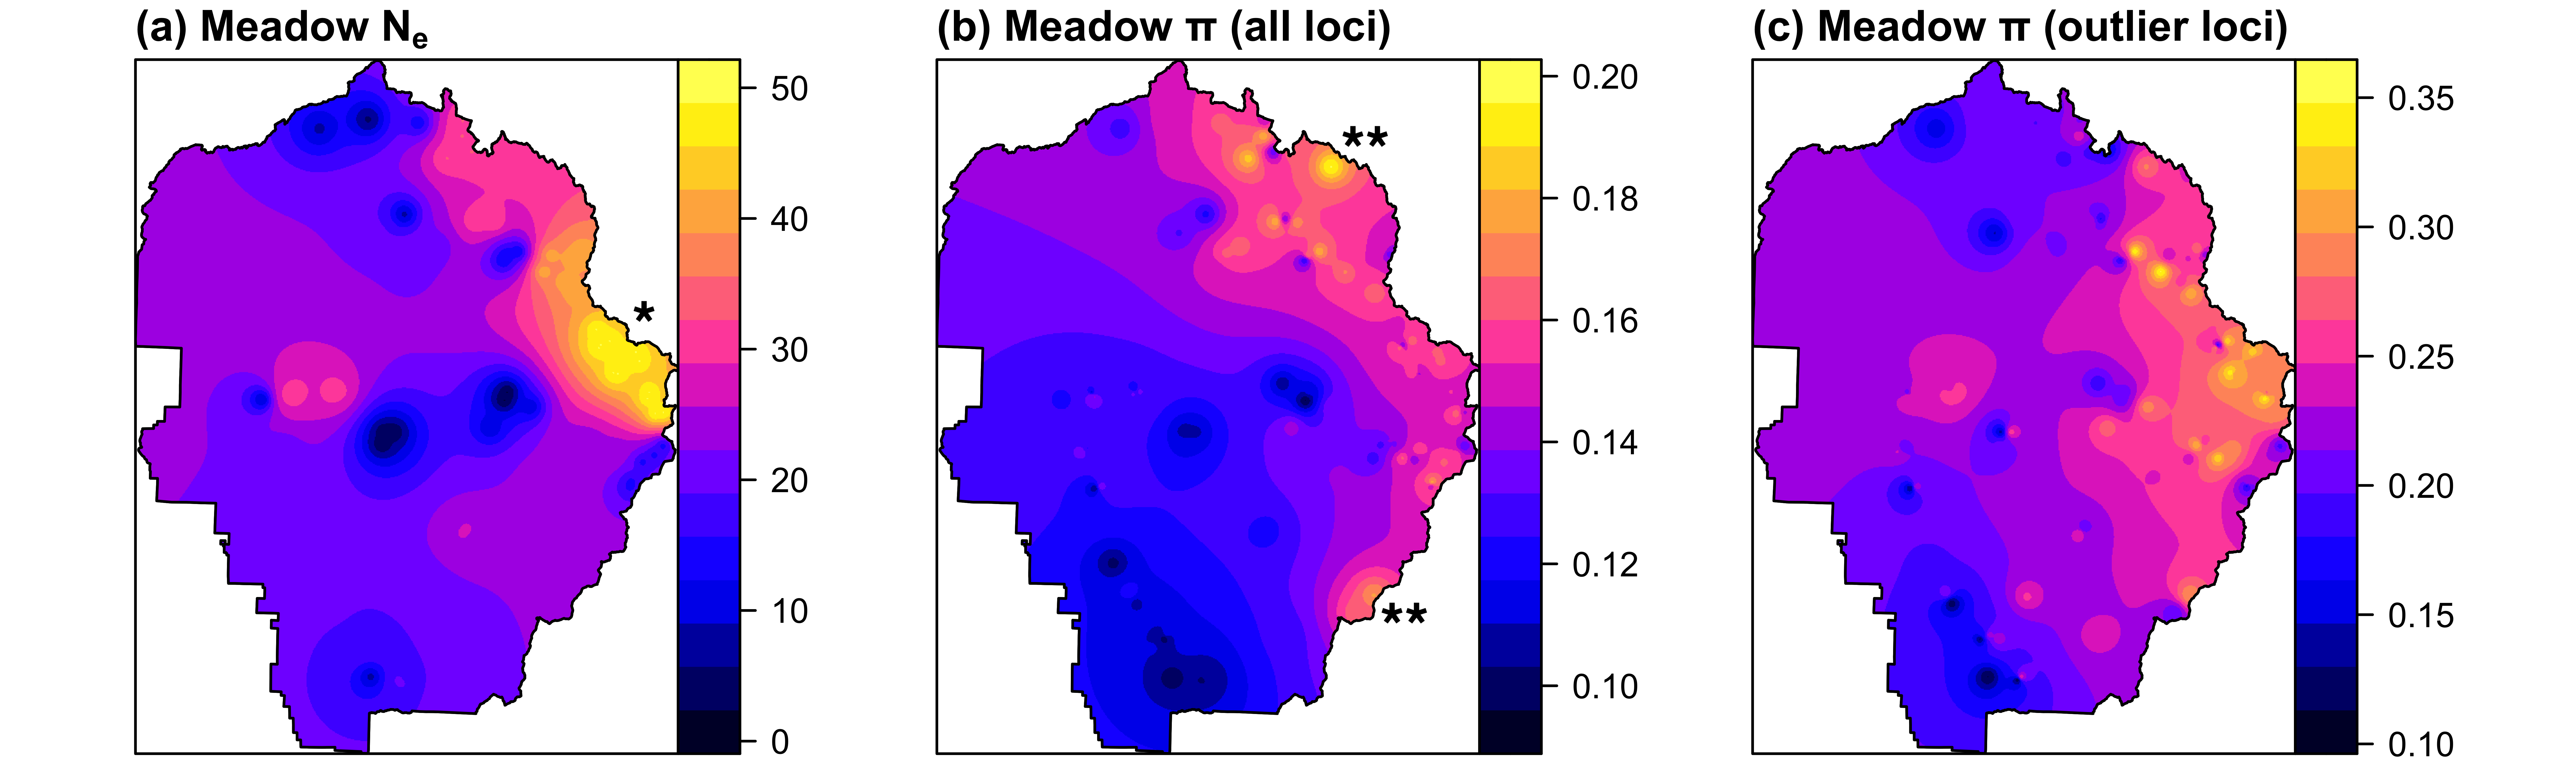
Working across roughly a third of all known Yosemite toad localities in Yosemite National Park, we mapped gene pool boundaries with recursive hierarchical clustering. The individual meadow turned out to be the most distinct unit, which is not a foregone conclusion for an animal that can walk. Average migration between meadows sits well under 10%, and the number of effective migrants per generation is well under one. Toad-occupied meadows in the park have a mean radius of about 266 m, so these are small, genetically distinguishable patches.
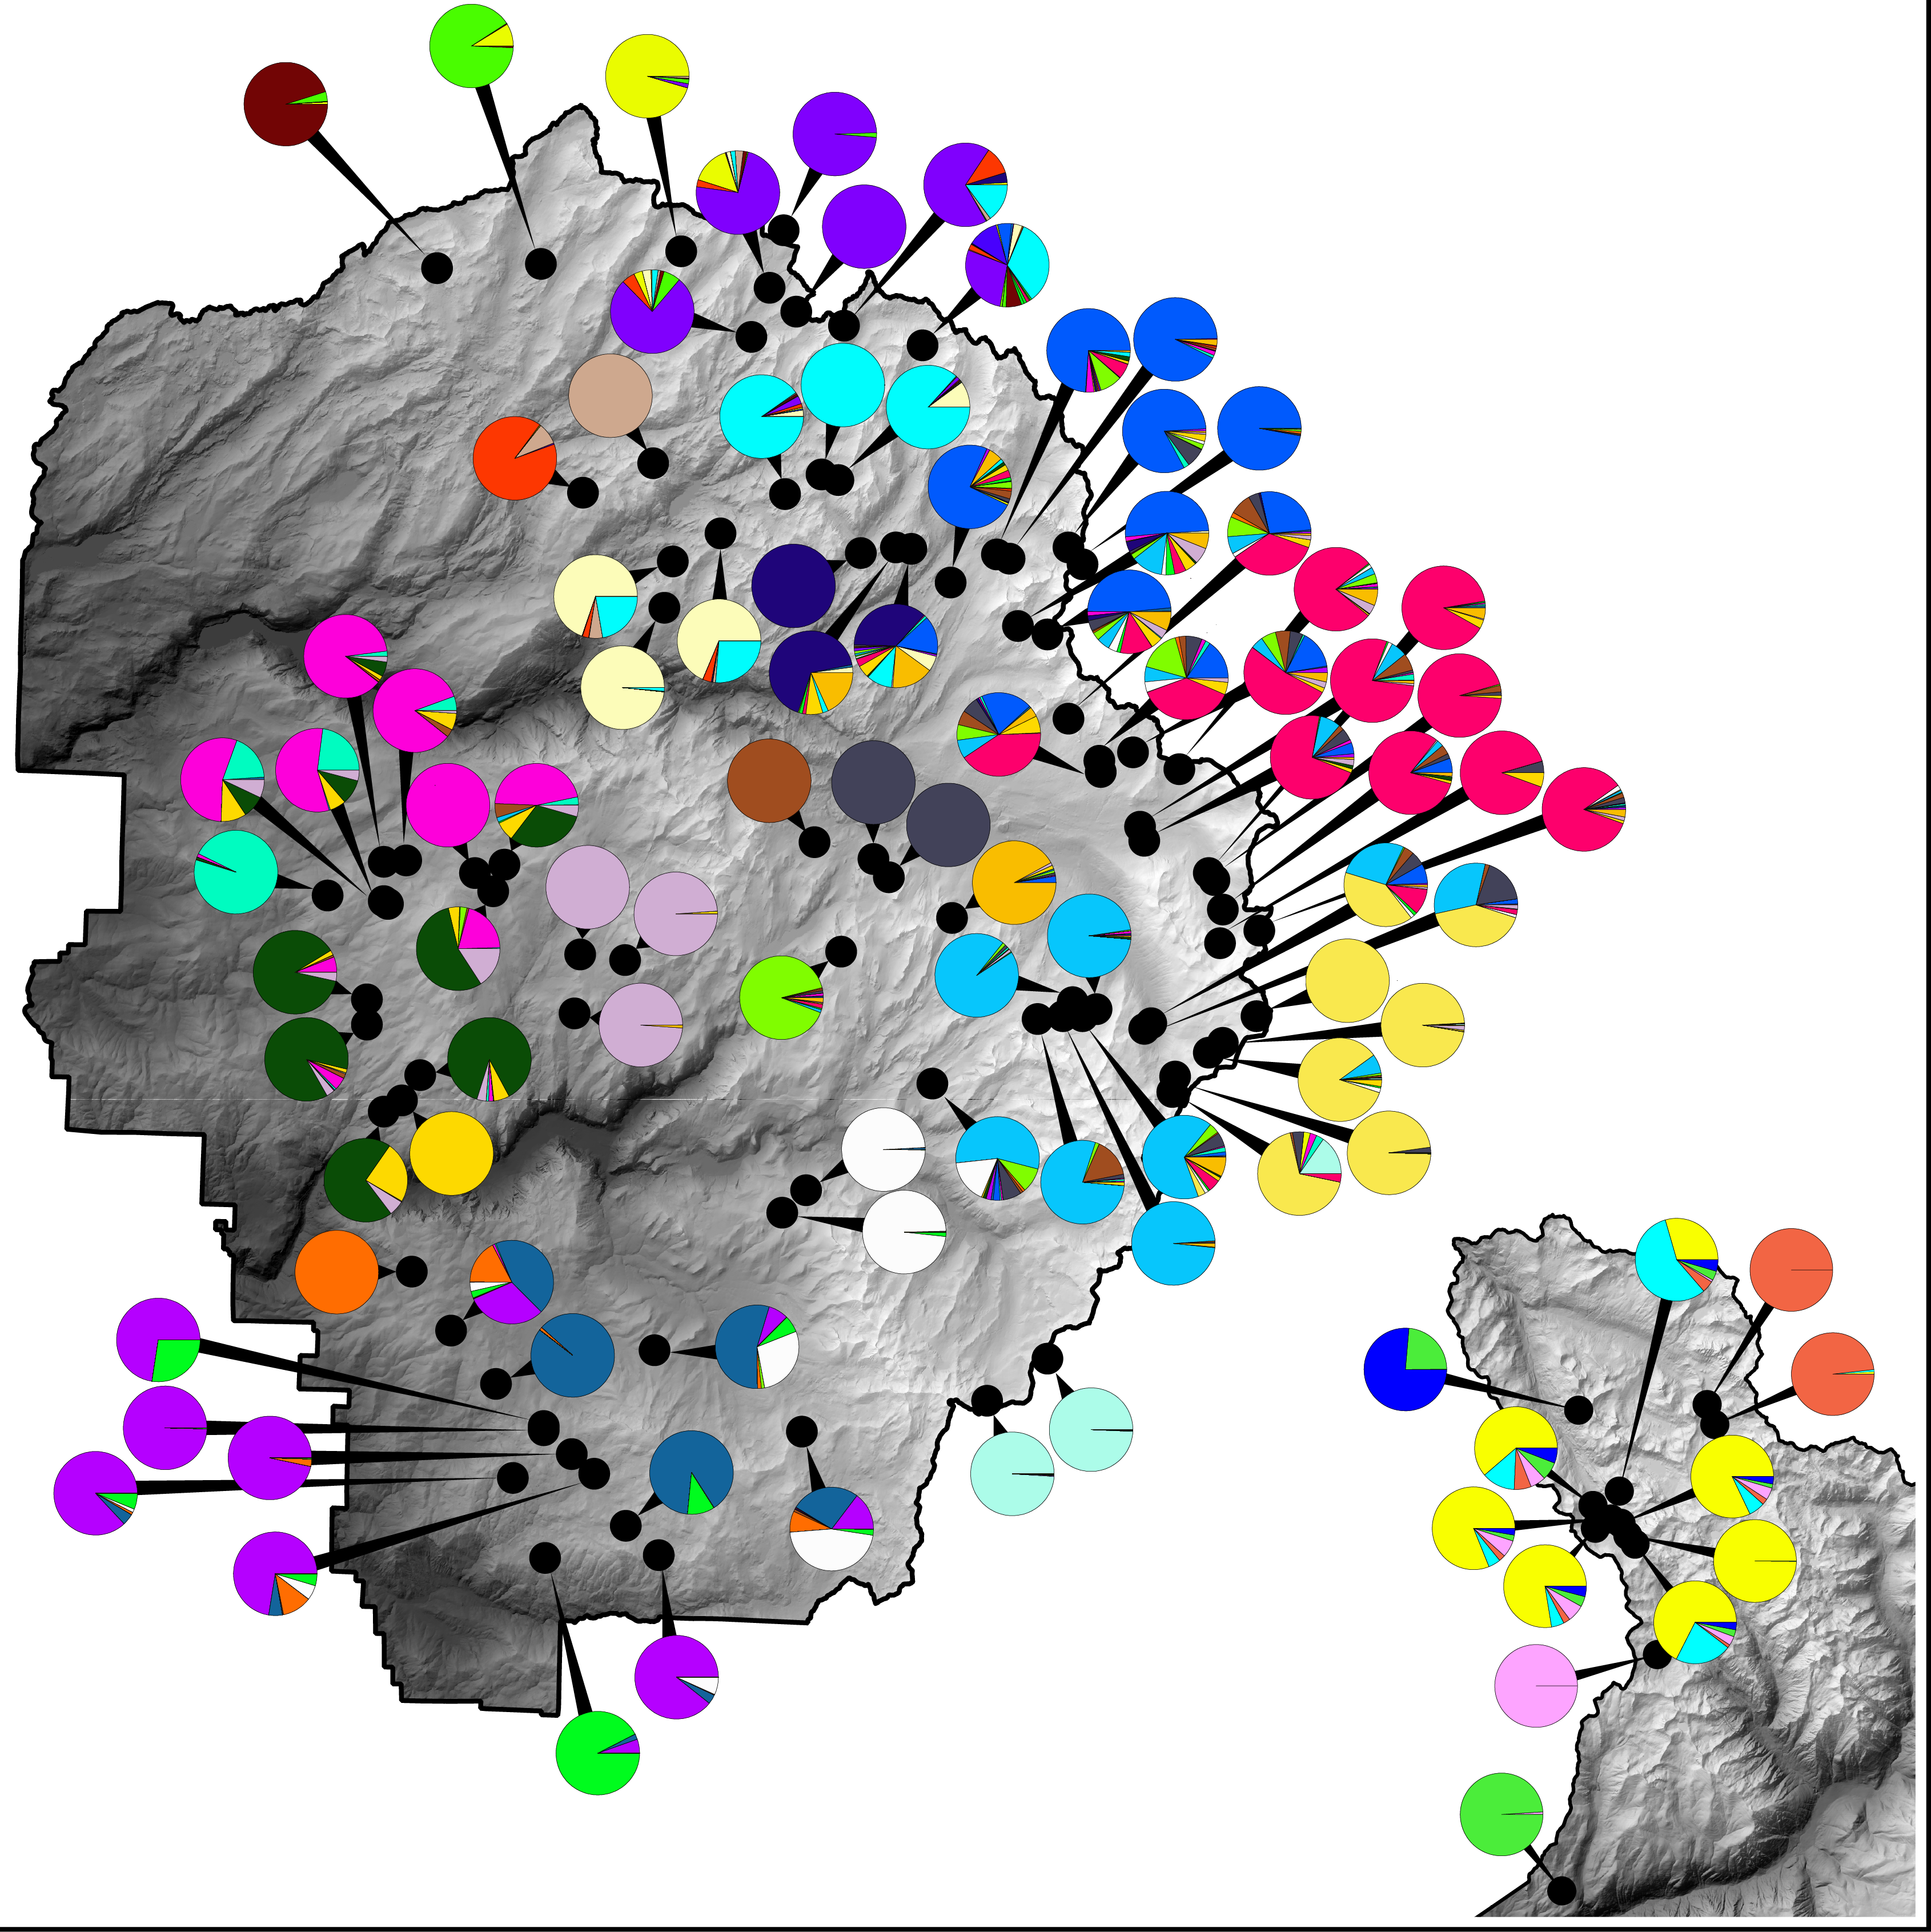
Meadows group into neighborhoods with substantially more movement inside a neighborhood than between. Within those neighborhoods, migration runs preferentially in one direction.
Neighborhoods often contain one or more large, flat hub meadows that experience net immigration, surrounded by smaller and topographically rugged satellite meadows with net emigration. Hubs also tend to hold more genetic diversity. That combination makes them obvious candidates for prioritized protection and as source populations for any reestablishment effort.
Technical details
Hierarchical genetic structure was resolved by recursive STRUCTURE runs down to the
smallest supported unit, across five hierarchical levels using 3,261 markers in 102 populations, a
substantial increase in sample size, marker count, and depth over the previous analysis of this system
(28 populations, a single marker, three levels).
Migration rates and their asymmetry were estimated per meadow pair, then summarized as net immigration and emigration to classify hubs and satellites. Migration estimates rest on island-model assumptions, but the headline result, that meadows are the most distinct unit, is robust to relaxing them.
Landscape genetics and the corridors that are moving uphill
Landscape genetics asks how habitat, topography, and climate shape connectivity between populations. Genome-wide differentiation among populations is produced by four processes: limited dispersal, environmental constraints on dispersal, incompatible adaptations among populations, and founder effects. Landscape genetic studies usually focus on the first two and implicitly assume only the present-day environment matters, leaving the other two unaccounted for.
The published approach works differently from the one in my dissertation. Rather than reducing the landscape to a single resistance surface, it identifies the most likely corridor between each pair of meadows and then optimizes the bandwidth of environmental influence around that corridor, because toads plausibly take multiple routes between any two meadows. This is analogous to circuit theory, except that paths and environmental influence are modeled separately, which keeps the raw environmental features intact instead of collapsing them before the analysis begins.
Snowpack-related features dominated: runoff, groundwater recharge, and related hydrological variables carried the most modeling importance. Under projected climate change the corridors shift upward and the asymmetry intensifies, meaning net movement increasingly runs uphill. That is a coherent response for a species tracking suitable habitat. It is also a problem, because there is a finite amount of uphill.
Technical details
Corridors were identified with least cost path analysis, then six candidate bandwidth types were evaluated per corridor. A random forests model trained on each of nine feature groups chose the optimal bandwidth independently for each group, so climate is free to act over a broader spatial scale than, say, geology.
Remote sensing inputs came from sources including LANDSAT and the Basin Characterization Model.
Features were reduced with PCA within groups and de-duplicated across groups.
Cubist regression trees then predicted and forecast both FST and the migration
asymmetry index δM, with permuted variable importance used to rank predictors. A
LineageCross term carried phylogeographic structure from the lineage analysis into the
connectivity model, which is how processes three and four above get accounted for.
Genomic geography: islands and rivers of speciation
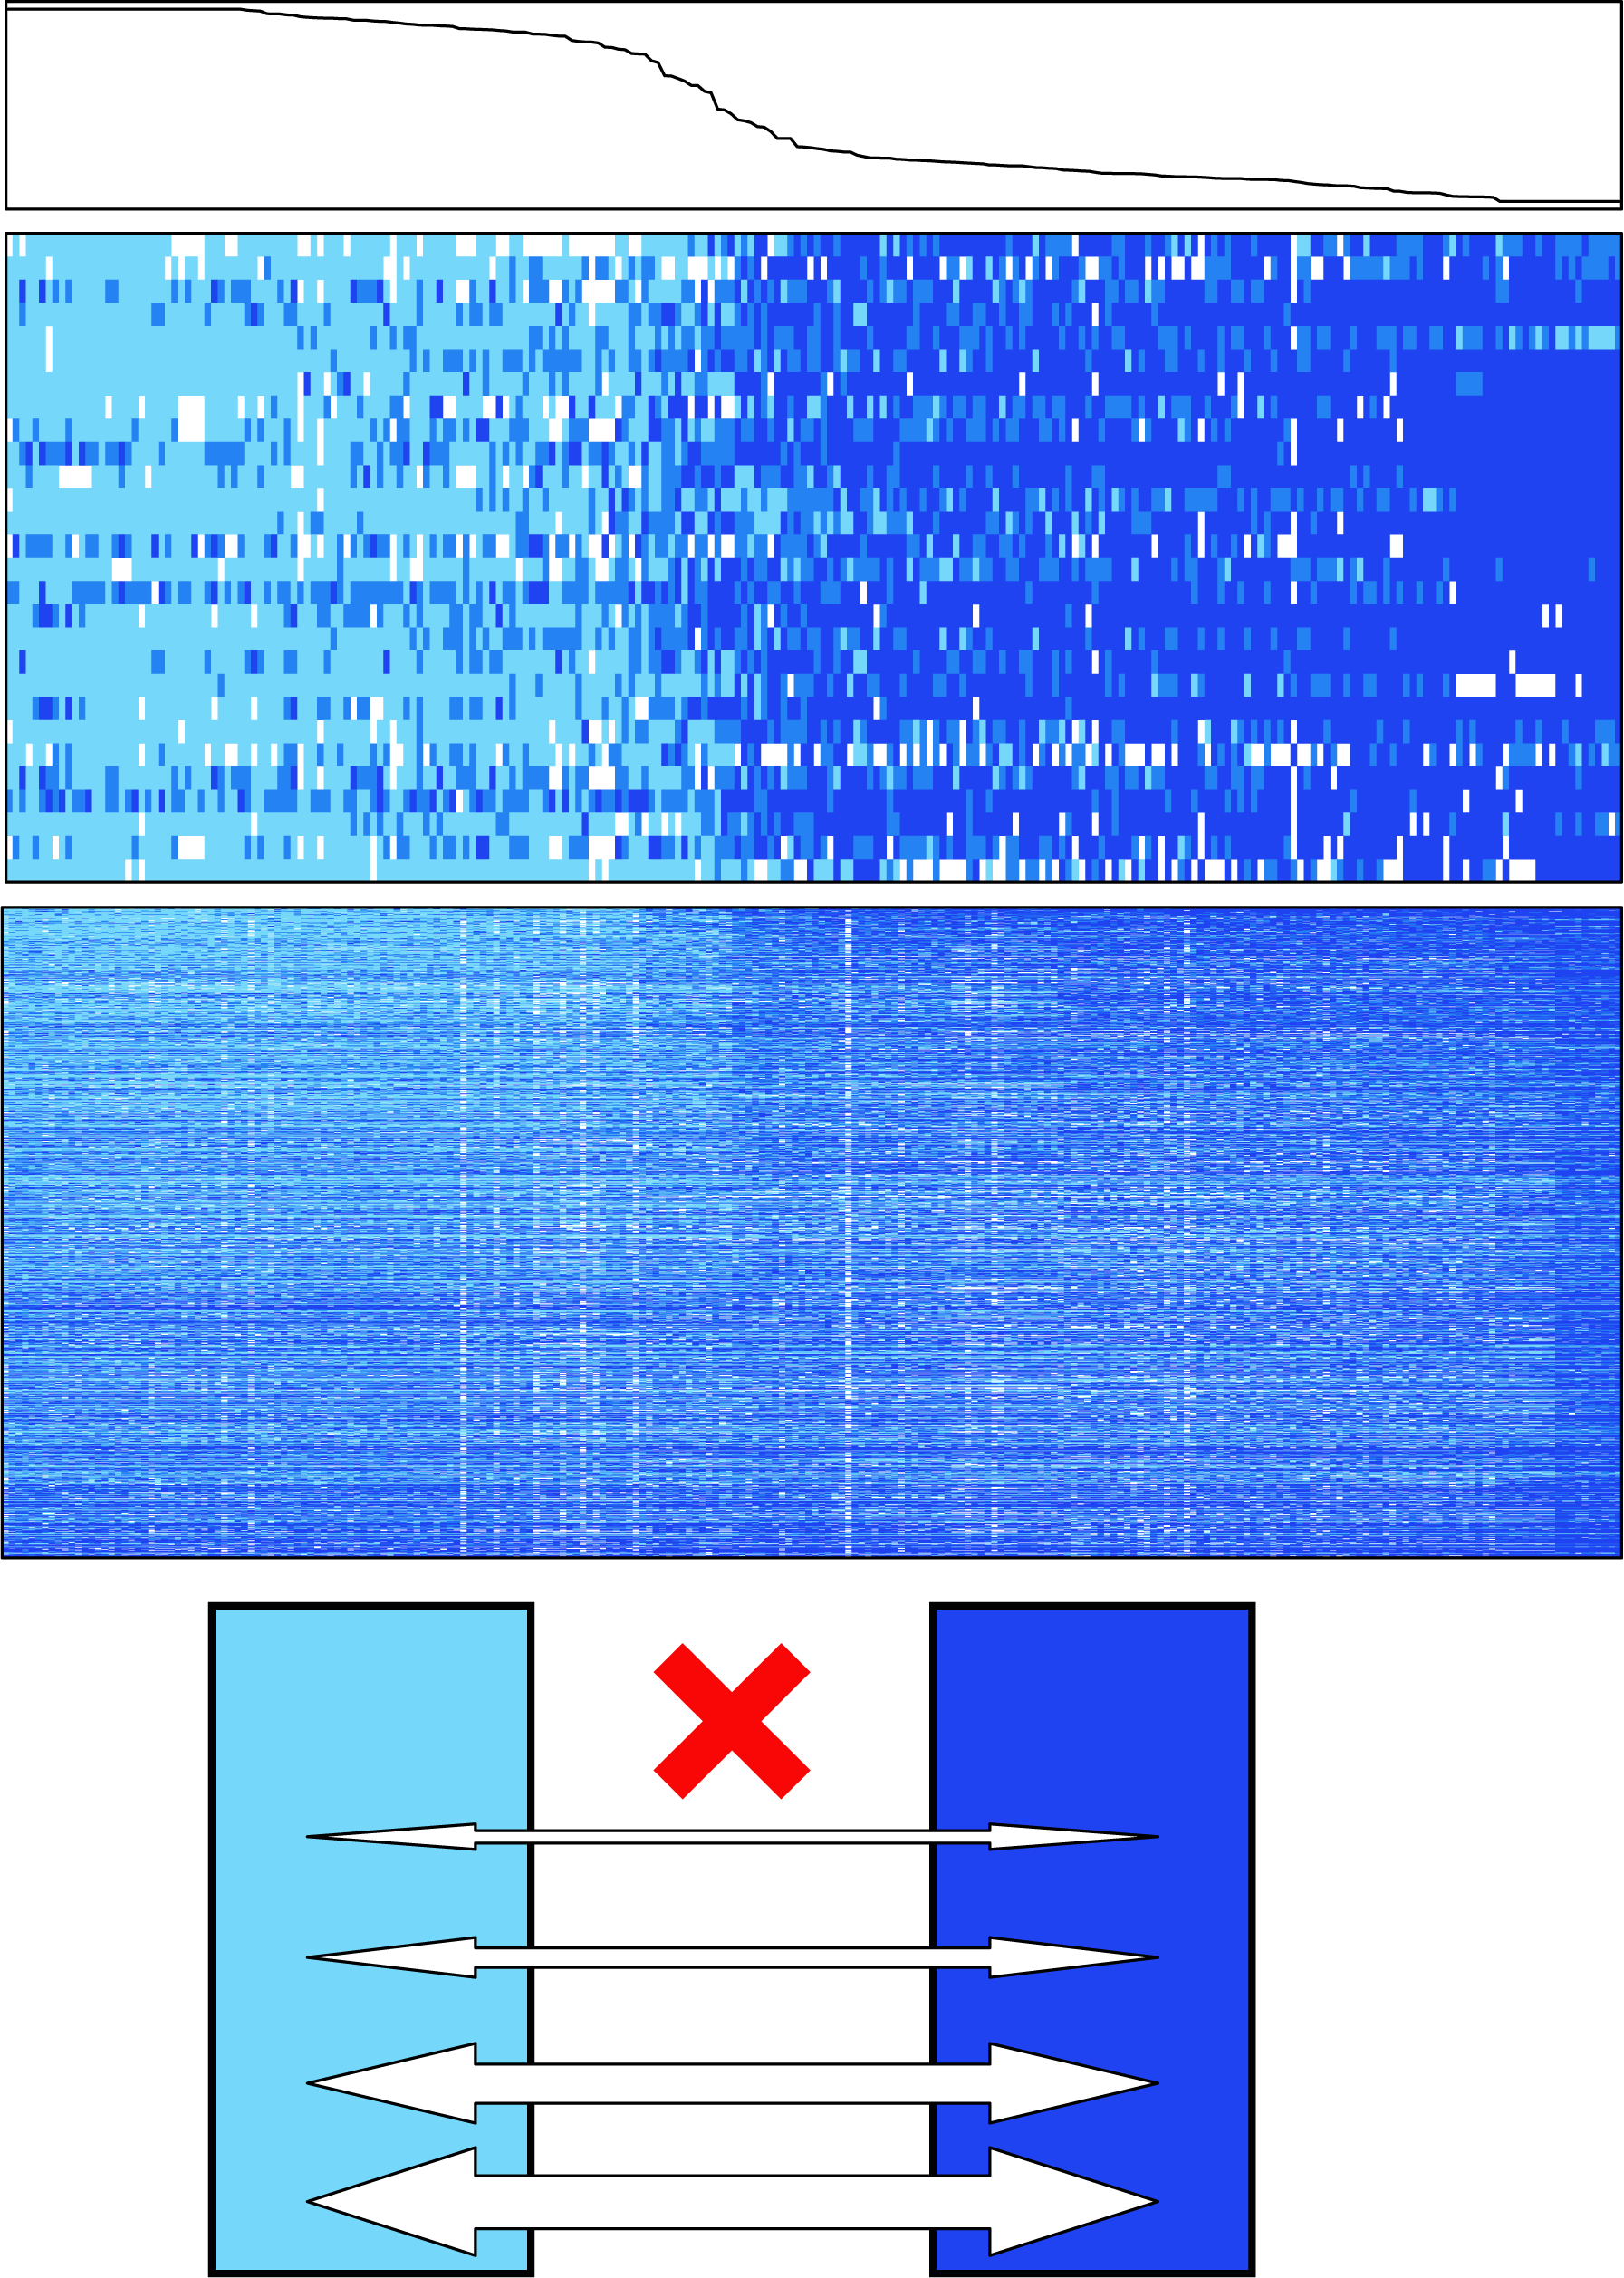
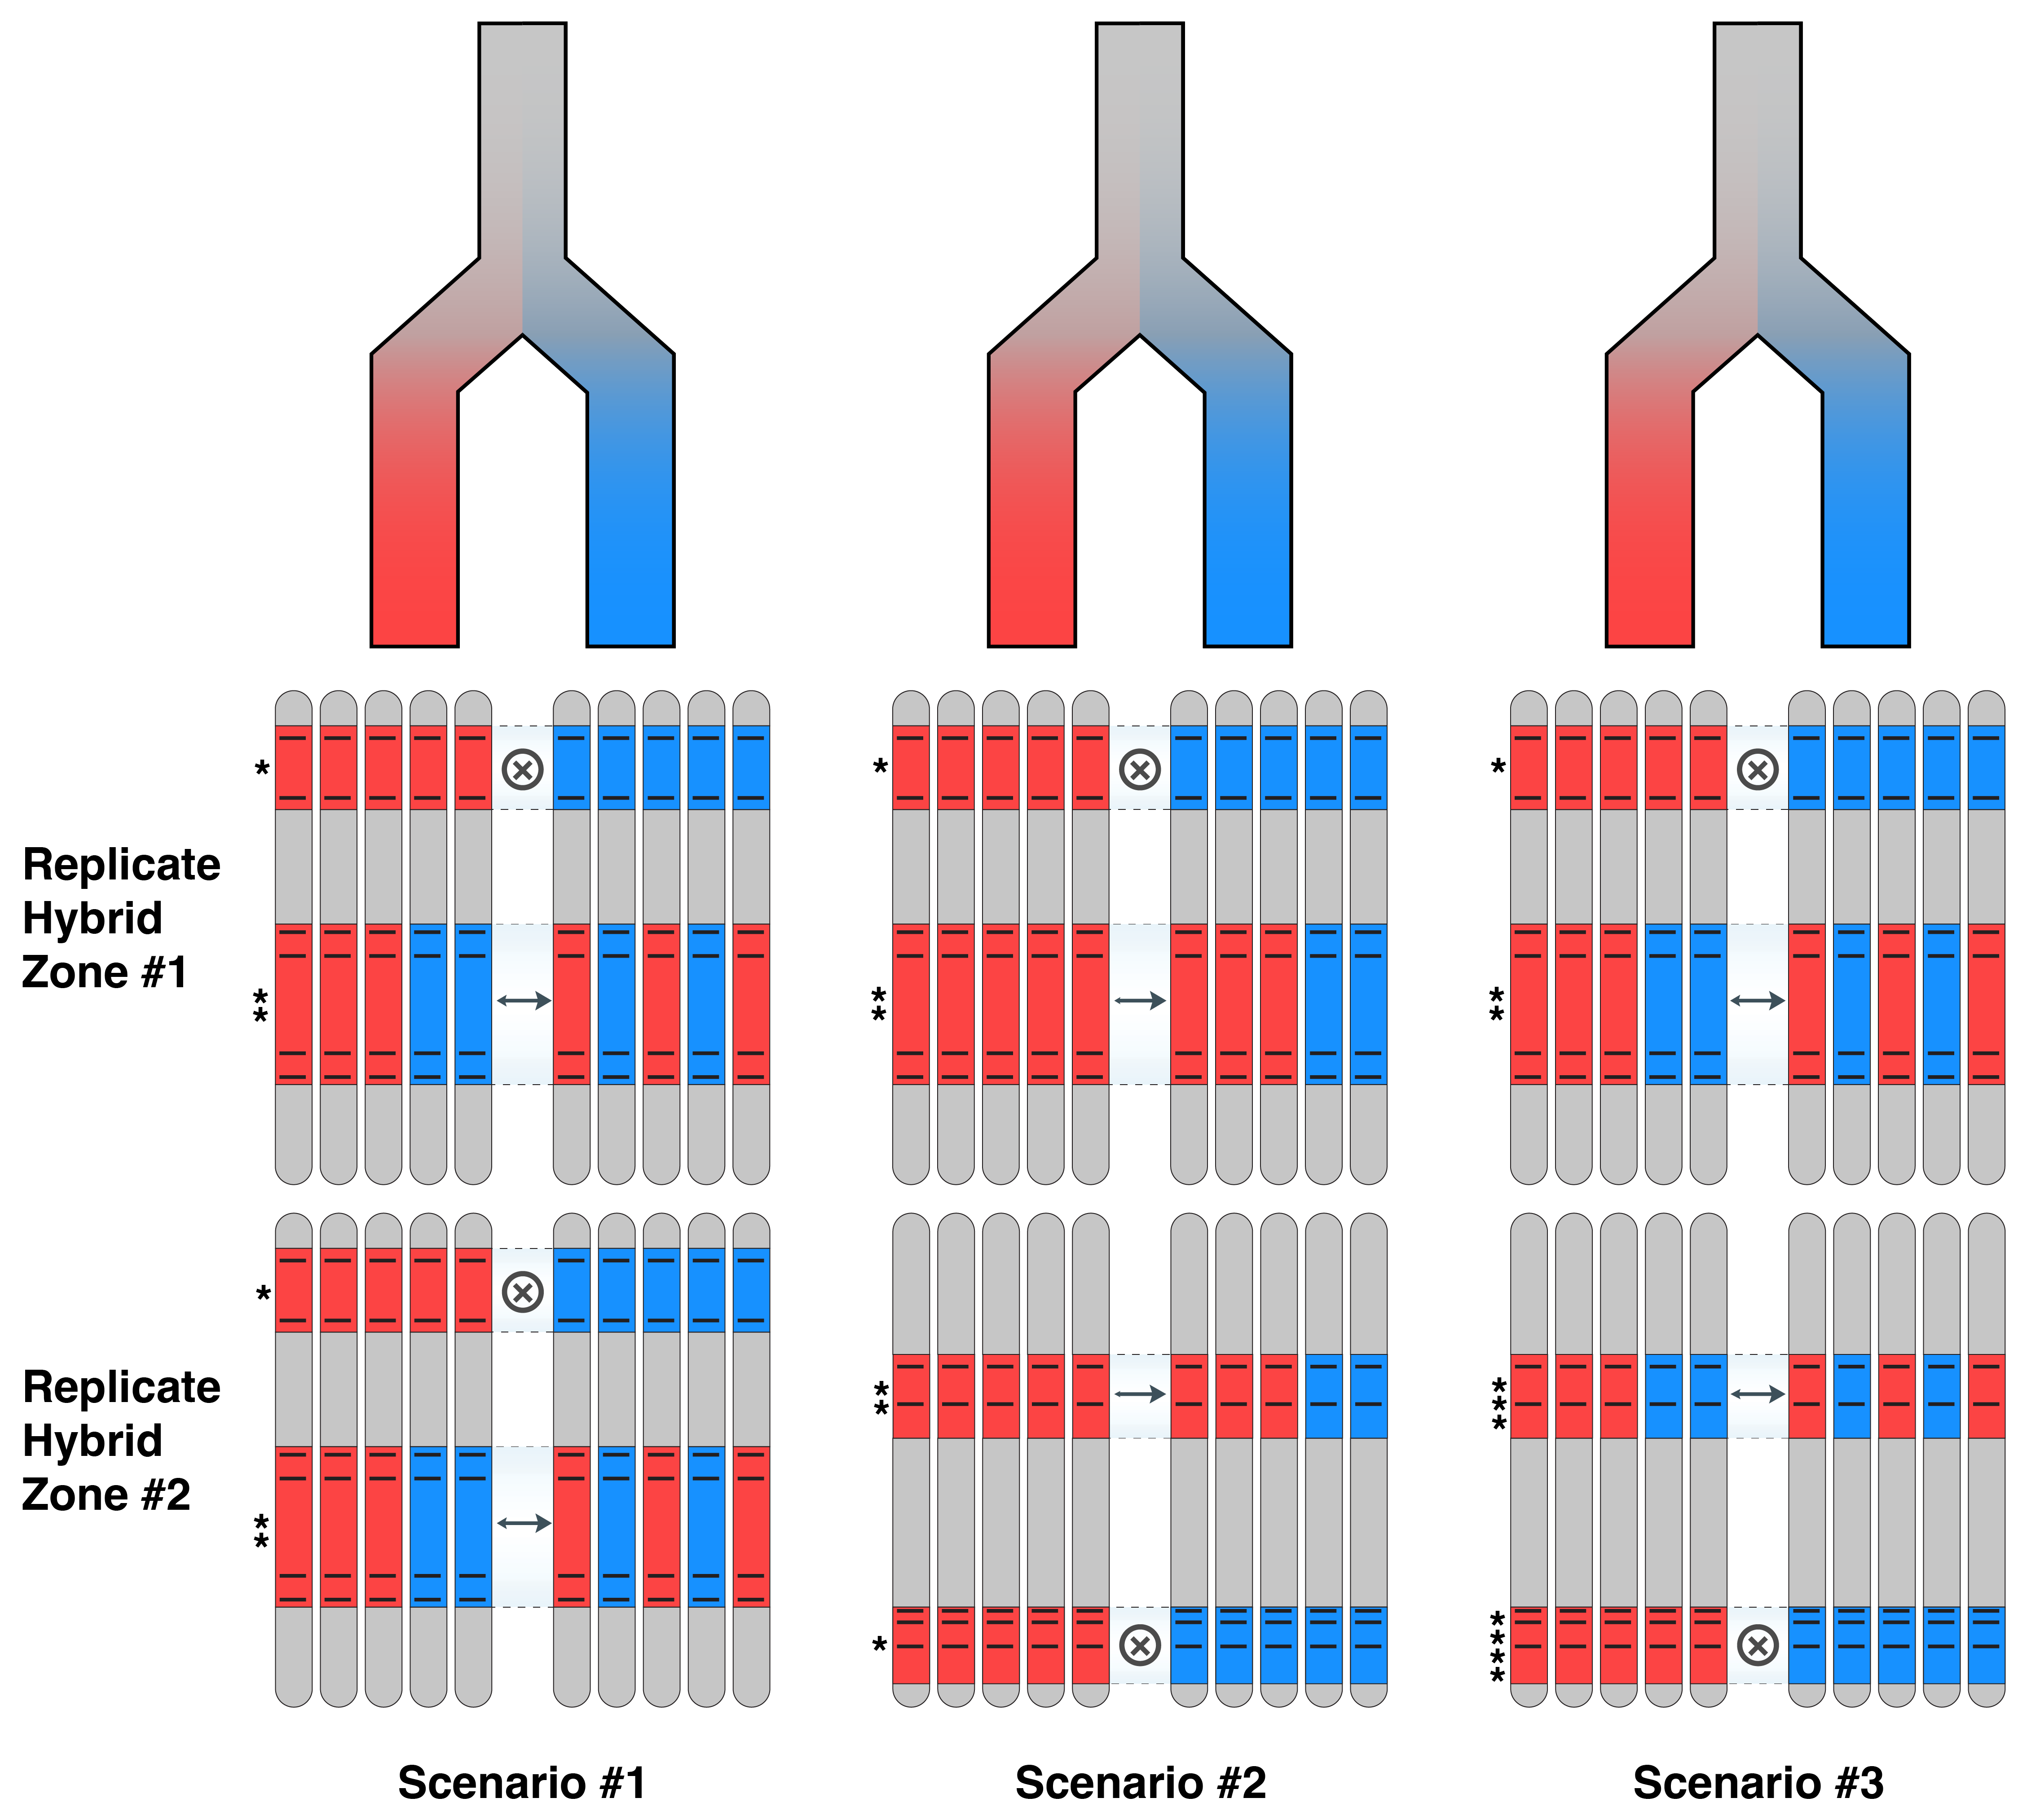
Hybridization and admixture between incipient species have traditionally been treated as destructive or as a nuisance. The reasoning is that some genes have fixed for alternate alleles, and hybrid combinations of those may be disadvantageous. These genomic islands are the exceptions to genome-wide patterns of gene flow, and they are often tied to the speciation process, as with color pattern genes in Heliconius butterflies.
Some genes under divergent selection tell the opposite story. They confer a benefit on hybrids and flow freely across the contact zone. These genomic rivers wash beneficial alleles into new genomic contexts, a process known as adaptive introgression, as with beak size genes in certain Darwin’s finches.
Very little was known about the gene identity of islands and rivers, whether they overlap, or whether the pattern repeats across contact zones. A neutral island could secondarily become an adaptive river. Using the three contact zones identified in the lineage work and the ddRADseq dataset, I compared the two marker types directly, then built a de novo larval transcriptome to match RAD markers to genes and ask whether islands and rivers differ in gene ontology, in SNP types, and in the severity of protein change.
The tadpole problem
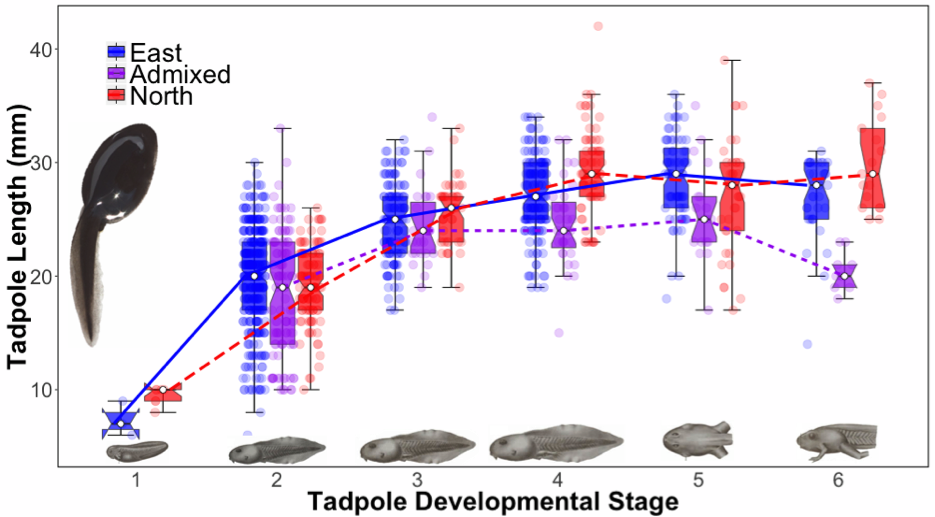
Tadpoles in a rapidly drying pond face a life history tradeoff: get big versus grow fast. Choosing wrong is the difference between metamorphosing and dying in the mud. Yosemite toads have a particular tendency to breed in very shallow pools, so desiccation and climate change bear directly on their biology.
There is phenotypic plasticity in this trait, letting an individual shift resources to match current pond conditions, but there is also a large genetic component. Couch’s spadefoot can metamorphose in a week. Several studies suggest genetic admixture specifically benefits individuals under pond desiccation, so I tested whether islands, rivers, or both relate to this trait.
The growth and development data are consistent with admixed larvae being better adapted to pond desiccation, and possibly to climate change. If that holds, it points toward a different paradigm for recovery planning in climate-threatened species. Under rapid environmental change, adaptation by mutation and selection may simply be too slow. When populations are isolated and then reunite, adaptive introgression of novel recombinant genotypes can move faster.
Technical details
Islands were identified with a multi-pronged outlier approach using hierarchical
ΦST, DXY, and the cladistic measure Slatkin and Maddison’s s.
Requiring a locus to pass all three tests gave stringent outlier counts of 14, 15, and 1 in the
east-north, east-south, and east-west contact zones respectively. Rivers were identified with Bayesian
genomic cline analysis in bgc, and asymmetrical gene flow was estimated per contact zone
with migrate-n.
The transcriptome used over 225 million reads from three tadpoles on an Illumina HiSeq 2500,
assembled and annotated with Trinity, with SNPs called and annotated through
GATK and Ensembl’s VEP. That yielded more than
517,000 transcriptome-wide SNPs.
The association analysis modeled overall growth-by-development with polynomial regression, then
fit each SNP in a linear mixed model using lmem.gwaser, with significant principal
components included as covariates to control for population structure. Gene ontology enrichment was
tested with Cochran-Mantel-Haenszel tests at level 2 of the GO hierarchy, checking independence of GO
terms by marker class while accounting for contact zone, and vice versa.
Delimiting conservation units for evolutionary potential
This is the chapter the rest of the work was building toward, and the piece of research I am most pleased with. Every earlier result feeds into it: the ancient lineage boundaries, the fused lineages and adaptive introgression at contact zones, and the current gene flow between meadows. On their own each answers a narrow question. Together they can be pointed at a much harder one.
Conservation units are the practical output of all this. They determine what gets protected, what gets counted as a distinct population, and where recovery money goes. Almost universally they are drawn from retrospective patterns: whole-genome measures of how isolated populations have been from each other historically. Units that are sufficiently isolated are assumed to hold separate evolutionary potential.
The problem is a mismatch of tense. The question conservation actually asks is forward-looking: which populations will persist and adapt? Answering it with a backward-looking pattern assumes the past predicts the future, and under rapid climate change that assumption is doing a great deal of unexamined work. Worse, genome-wide isolation measures are dominated by neutral variation, which says a lot about drift and dispersal and little about the capacity to respond to selection.
Geminate Evolutionary Units
We proposed using both neutral and putatively adaptive variation to build explicitly predictive units, formalised as the Geminate Evolutionary Unit: a population expected to adapt similarly to models of future environmental change, and potentially to form a new lineage. The name borrows from geminate species, pairs that diverge after a barrier splits them, because that is what these units are positioned to become.
The framework has four steps, and each is a separate analysis:
1. Identify loci under environmental selection.
2. Model and map the adaptive units that could spawn future lineages.
3. Forecast the relative selection pressure each unit will face.
4. Estimate fitness and likelihood of persistence with geo-genomic simulations.
Applied to Yosemite National Park, this produced three GEUs with contrasting adaptive optima. YF-North occupies high winter snowpack with moderate summer rainfall. YF-East occupies low to moderate snowpack with high summer rainfall. YF-Low-Elevation occupies low snowpack and low rainfall.
The contact zones did not resolve cleanly into one unit or another, which is the honest outcome rather than a failure. They were assigned as intermediate and overlapping GEUs on the basis of mixed clustering and unusually high genome-wide diversity, on the reasoning that a zone carrying novel recombinant variation may matter more for future adaptation than its ambiguous membership suggests.
Supply and demand
The useful way to read the result is as a balance between two quantities. Supply is how much climate-associated genetic variation a unit already holds, which sets how fast it can respond. Demand is how much climatic change it will actually experience, which sets how much response is required. A unit fails when demand outruns supply.
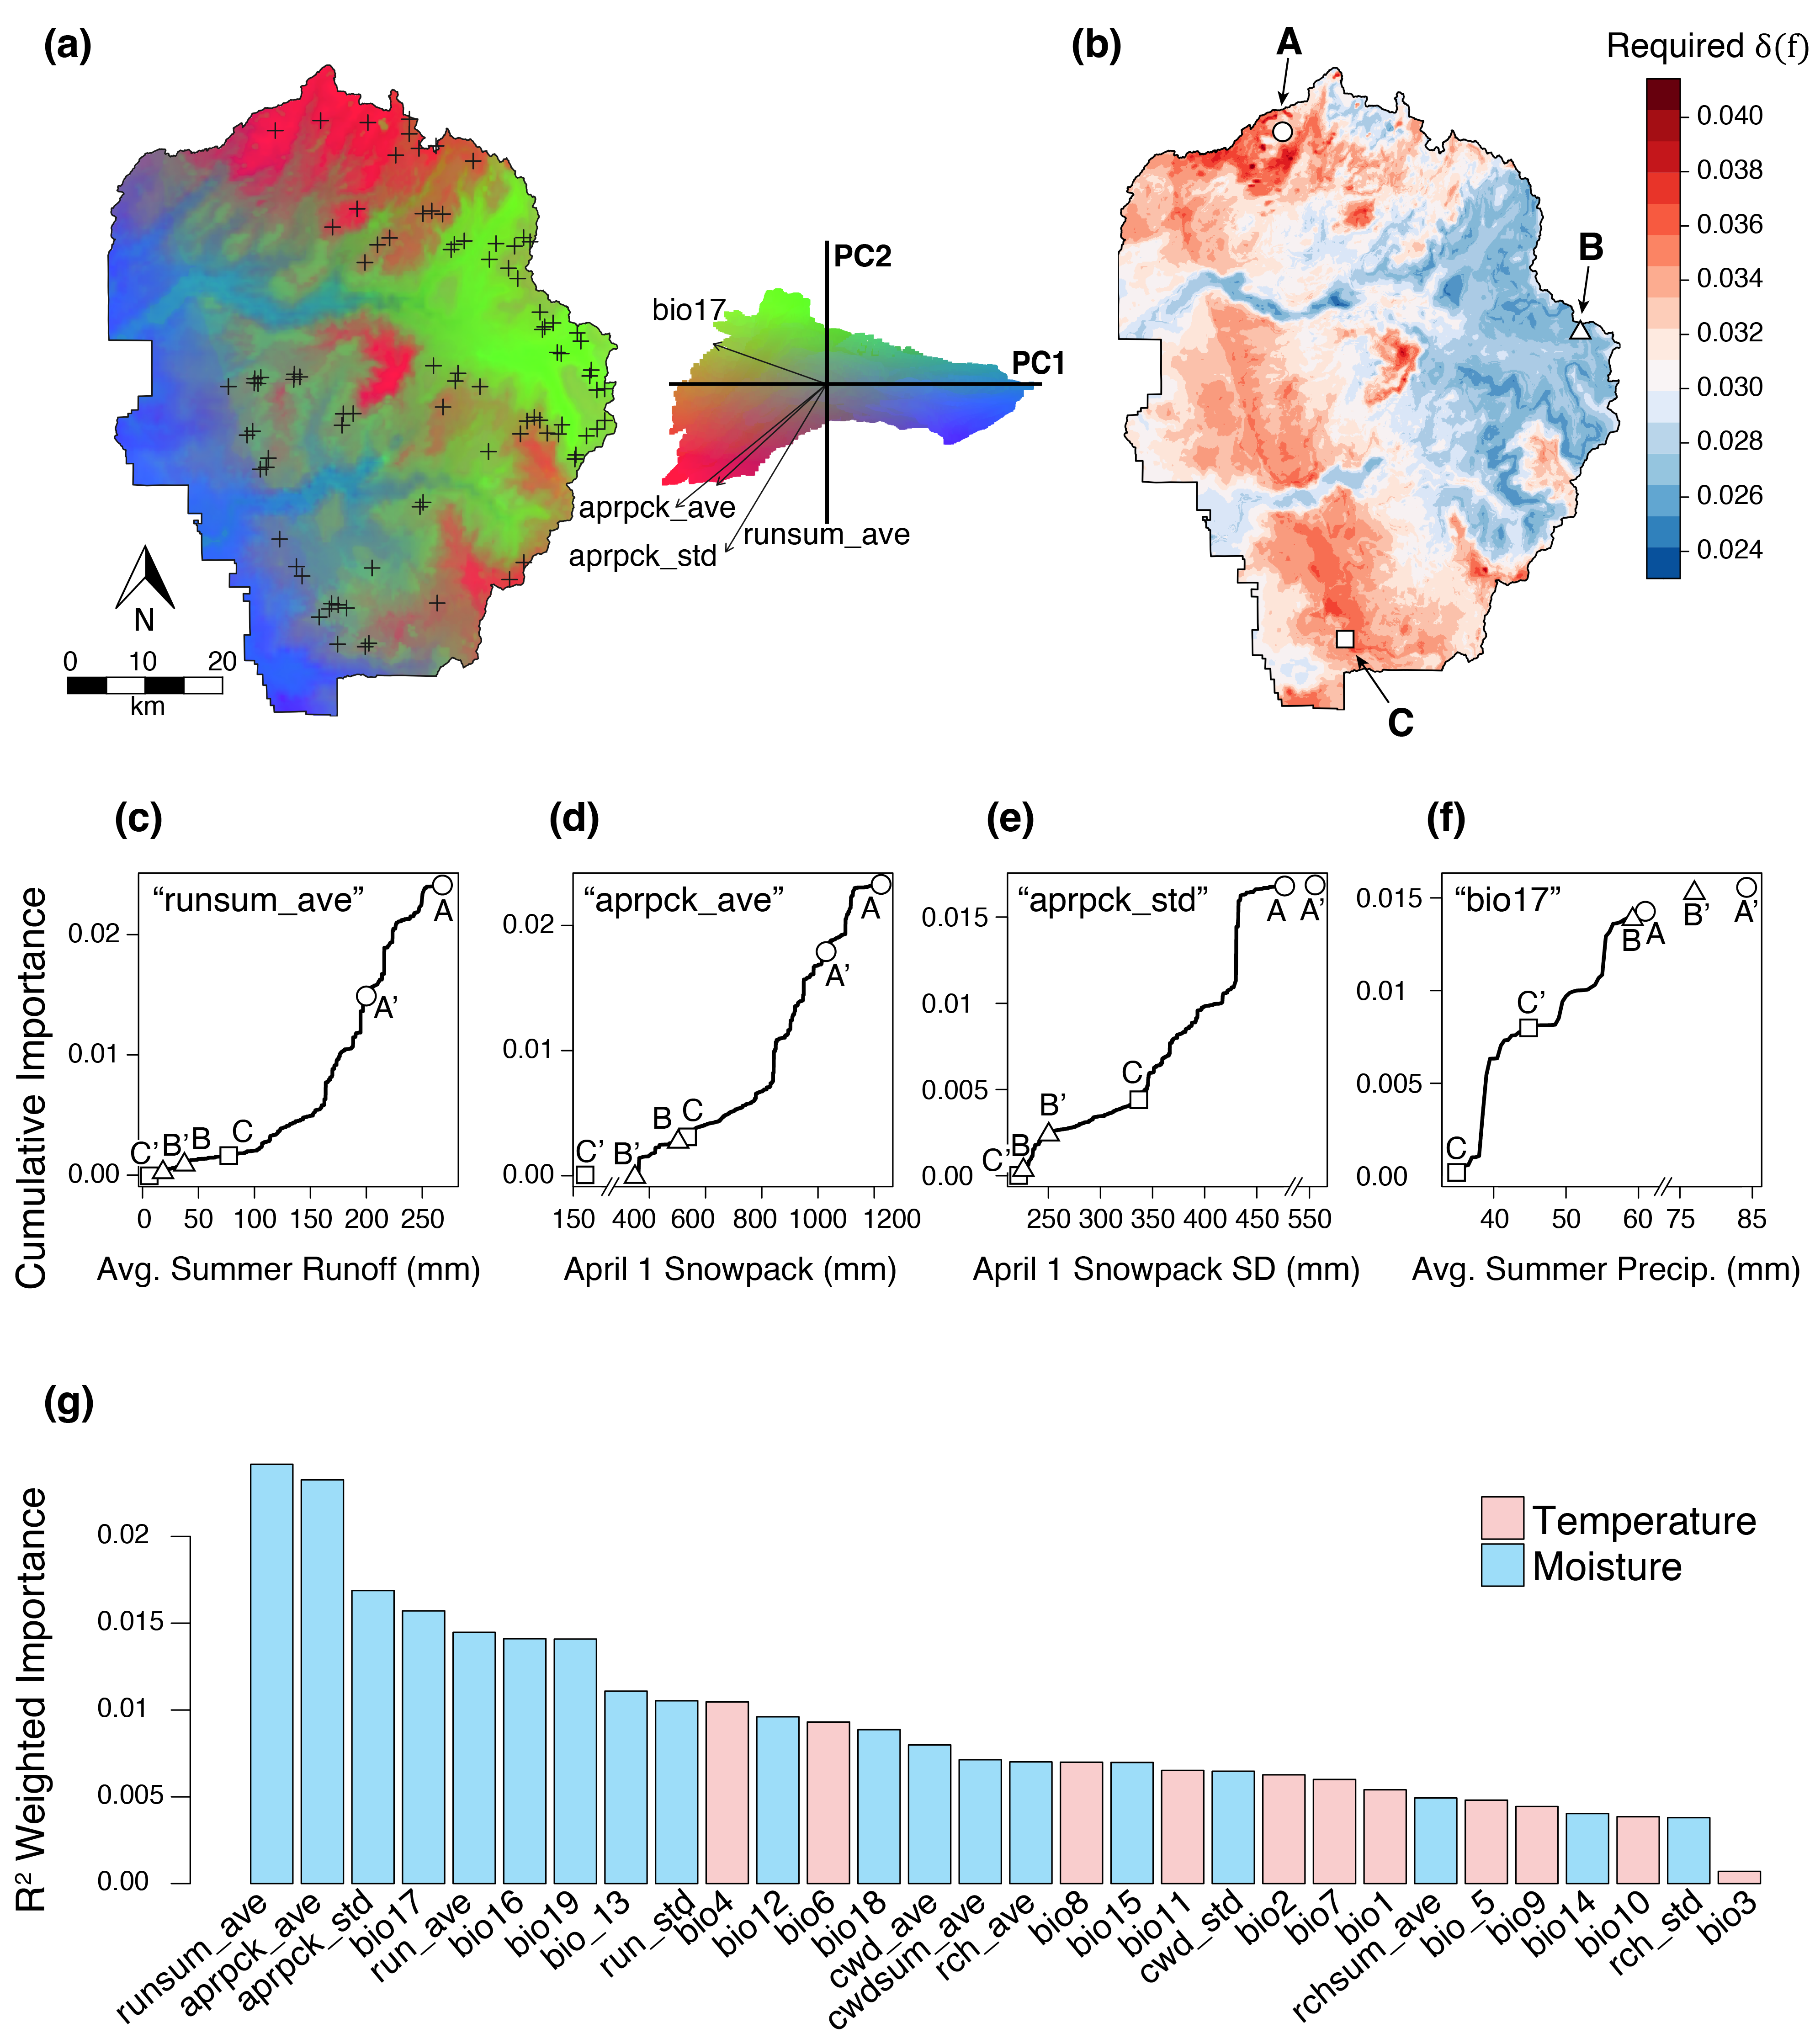
Separating the two is what the framework buys you. A population can be genetically impoverished and still do fine if it sits somewhere the climate barely shifts; a diverse population in a fast-changing place can still fail. Collapsing both into a single vulnerability score hides exactly the distinction a manager needs in order to choose between protecting a place and protecting a gene pool.
Simulating the outcome
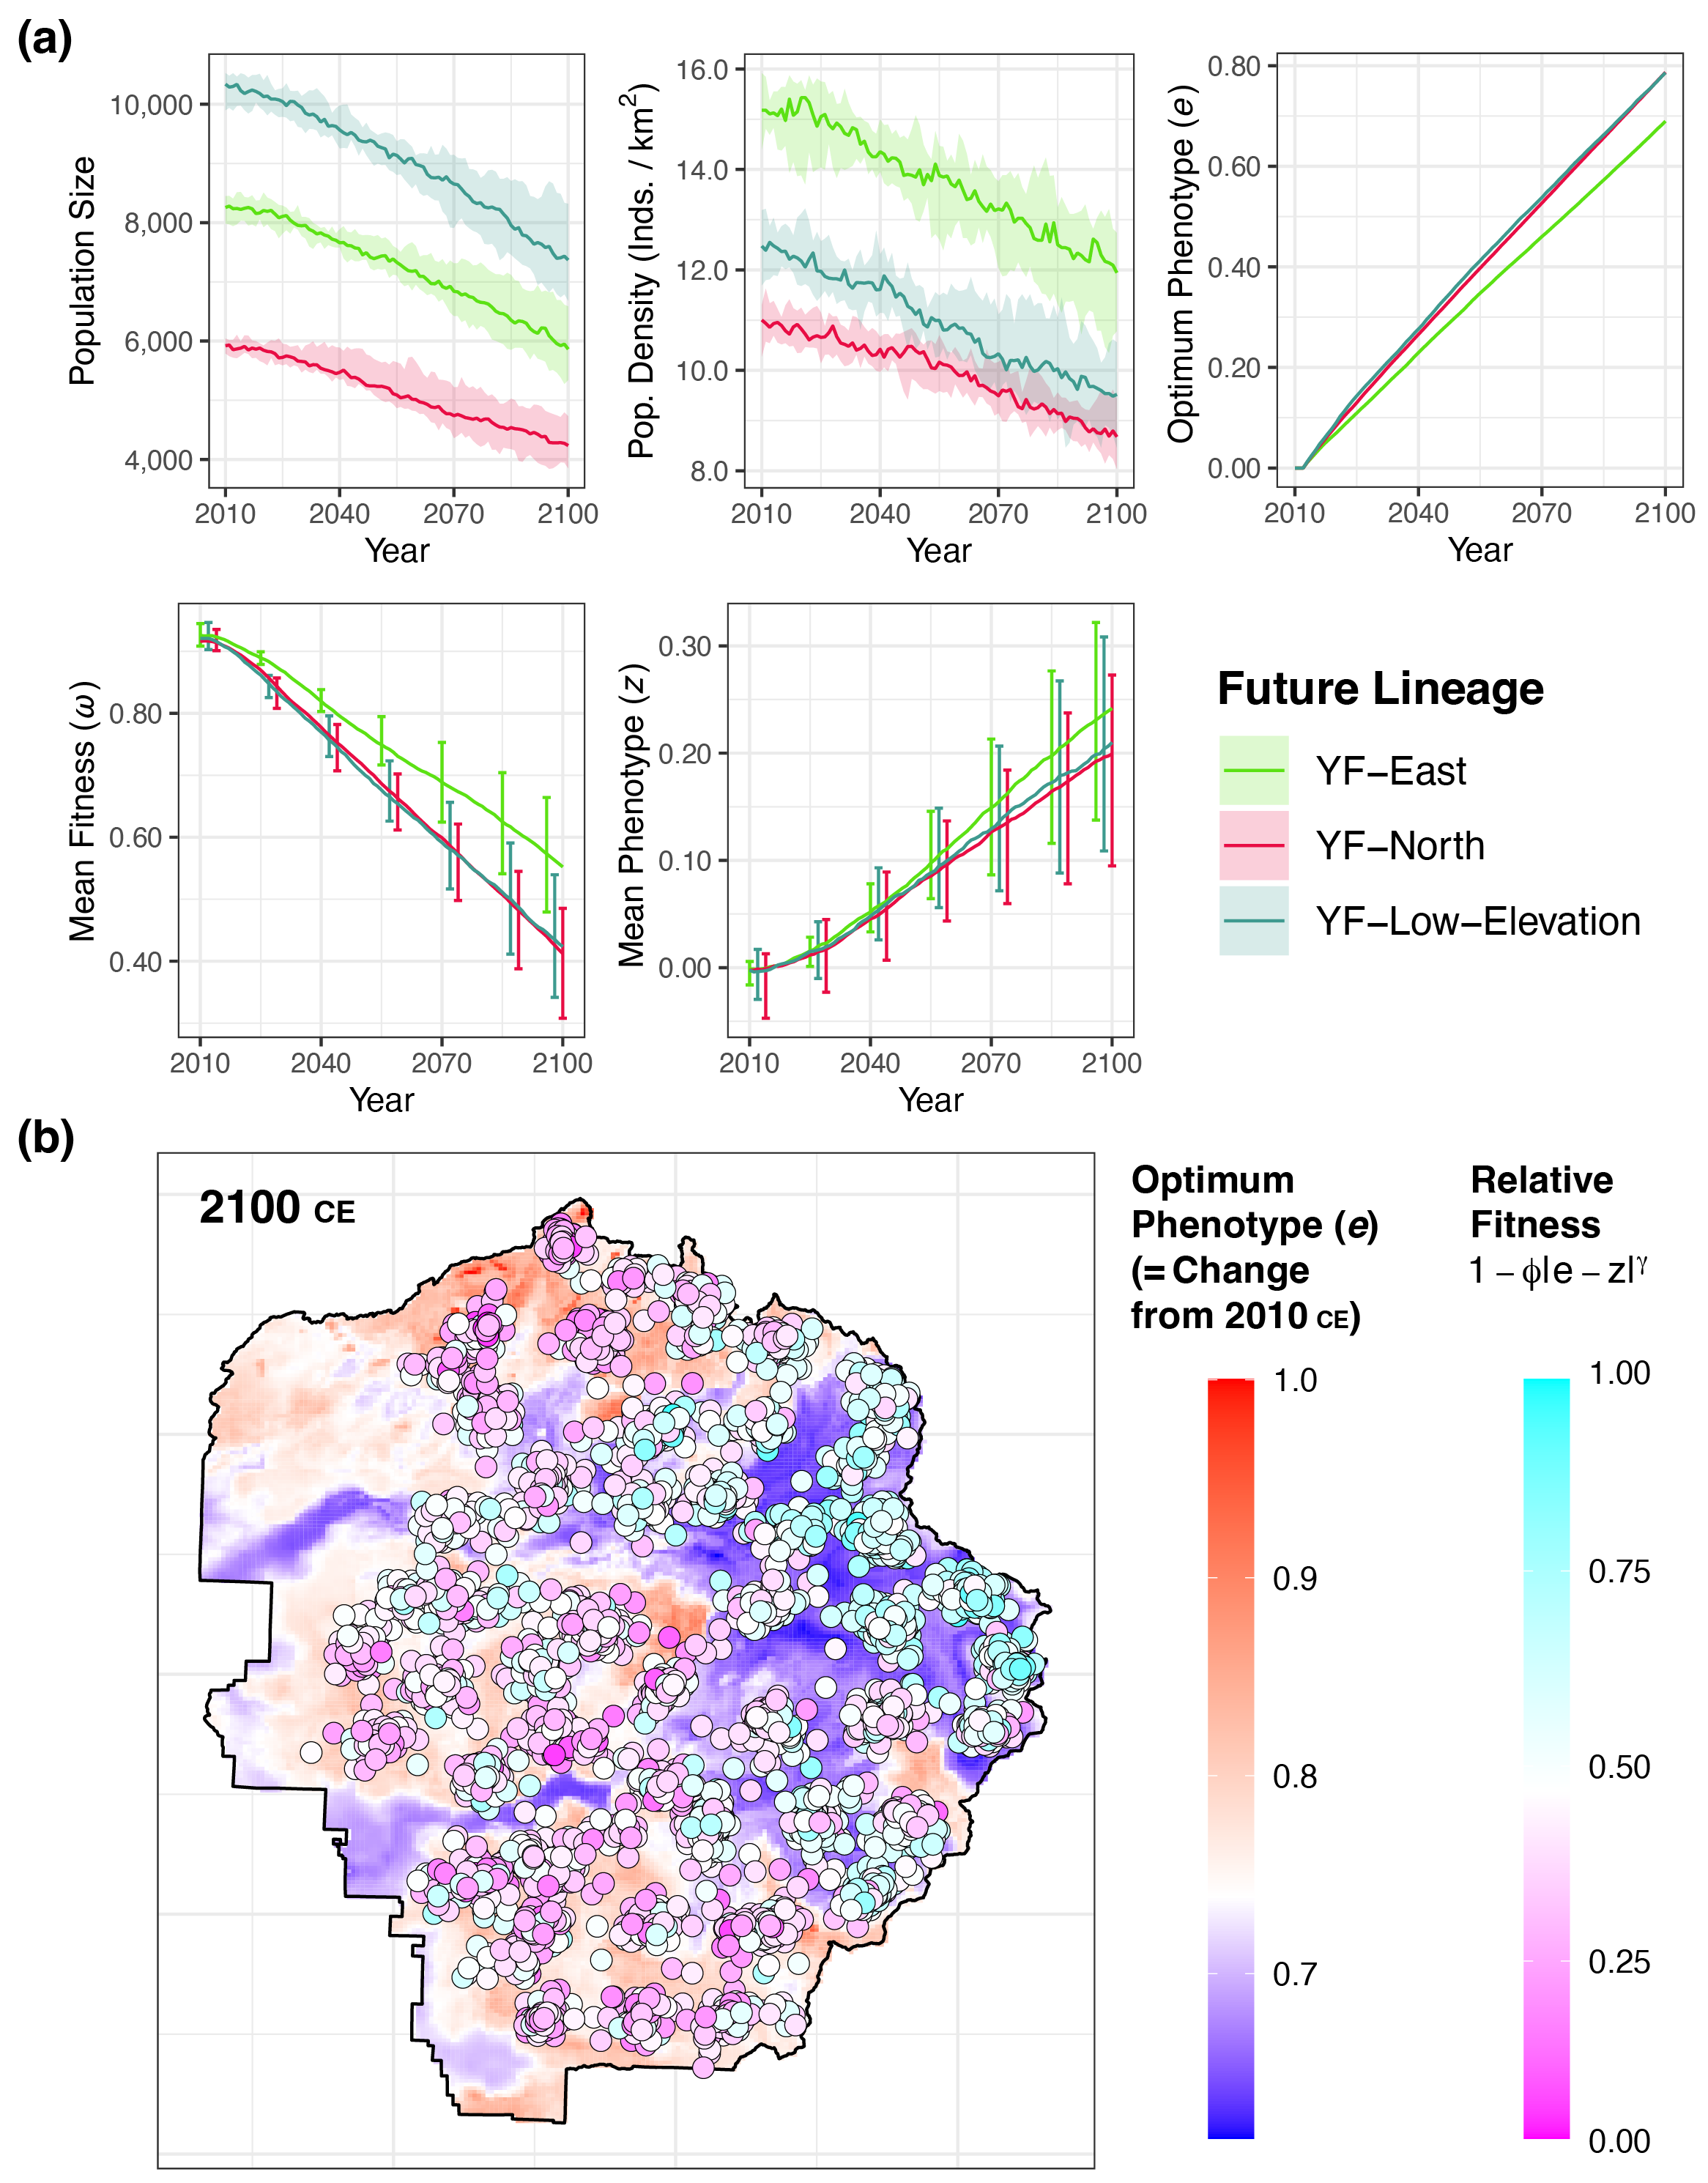
The last step converts a map of adaptive variation into a statement about survival. Geo-genomic simulations run each projected lineage forward under a climate scenario, tracking allele frequencies against shifting optima, and return relative fitness rather than a habitat suitability score.
Under RCP 8.5 the simulations project a 29 percent species-wide decline over 90 years. The most diverse unit, YF-East, is least affected, for two reasons that map onto supply and demand: it sits where climatic selection pressure will be weakest, and its standing genetic diversity supports the fastest adaptive response. Ranked from most to least vulnerable: YF-Low-Elevation, YF-North, YF-East.
A 29 percent decline in under a century is a startling number for a species already listed as threatened, and it is consistent with observed declines elsewhere in the Sierra Nevada. The framework itself is the transferable part. The specific loci belong to this toad; the logic of delineating units by projected adaptive capacity rather than historical pattern applies to any imperilled species facing rapid environmental change, and most of them are non-model organisms where this is the only kind of evidence available.
Technical details
Candidate loci under climatic selection were identified with redundancy analysis and Bayesian
association methods, controlling for demographic structure so that population history does not
masquerade as selection. That yielded 24 candidate loci with R2
from 0.09 to 0.52, including genes such as MAP3K5, which is involved in cellular response
to environmental change.
Future genomic response was forecast with Gradient Forests, a multivariate machine
learning method that maps allele frequency turnover onto environmental gradients and so allows the
required adaptive shift to be expressed in the same units as the available variation. Relative fitness
of each projected lineage came from geo-genomic simulations under RCP 8.5.
Part of this analysis is written up as a tutorial here. Data are archived at Dryad.
Hybridization with the western toad
In 1956, a biologist described a strange toad near Blue Lakes in Eldorado National Forest, 30 miles north of the Yosemite toad’s known range, and suggested it was a hybrid with the widespread western toad (Anaxyrus boreas). The claim was disputed, revisited, and never resolved. Surveys kept turning up intermediate-looking animals. Toads of uncertain species kept being killed on an off-highway vehicle trail, which matters legally, because one parent species is federally protected and hybrids have ambiguous status under the Endangered Species Act.
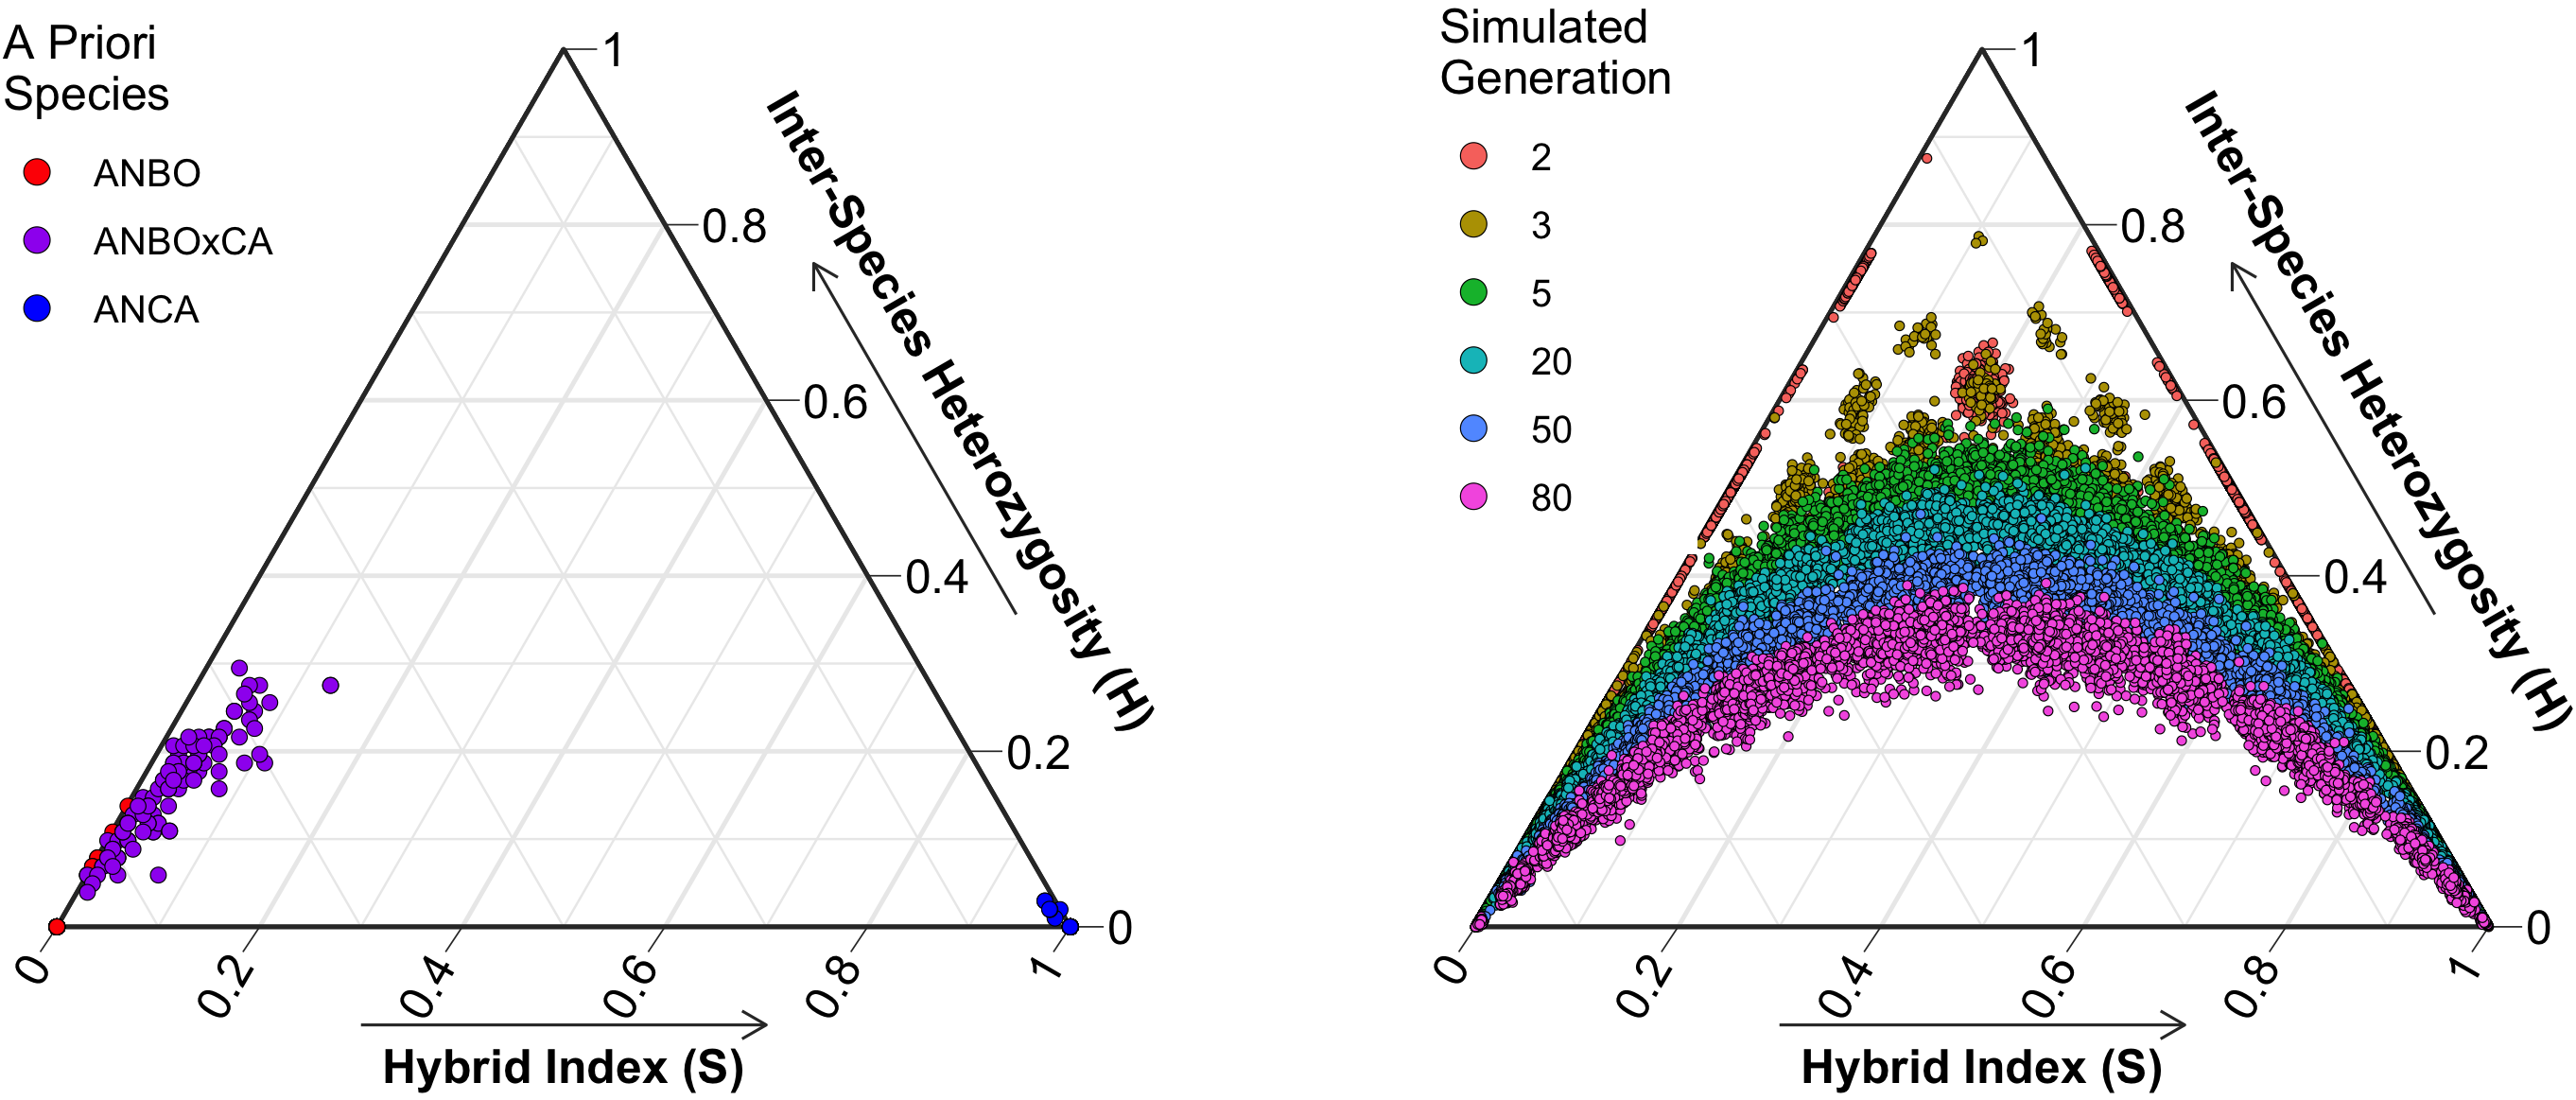
Morphology cannot settle this. Hybrid traits are not reliably intermediate because of dominance and epistasis, no two hybrids are alike after recombination, and several generations past the original cross an animal can look entirely like one parent. Identifying a fifth-generation backcross with 95% confidence takes about 95 loci; a tenth-generation backcross takes over 3,000. So we built a genomic hybrid panel, merging new samples with the existing Yosemite toad dataset.
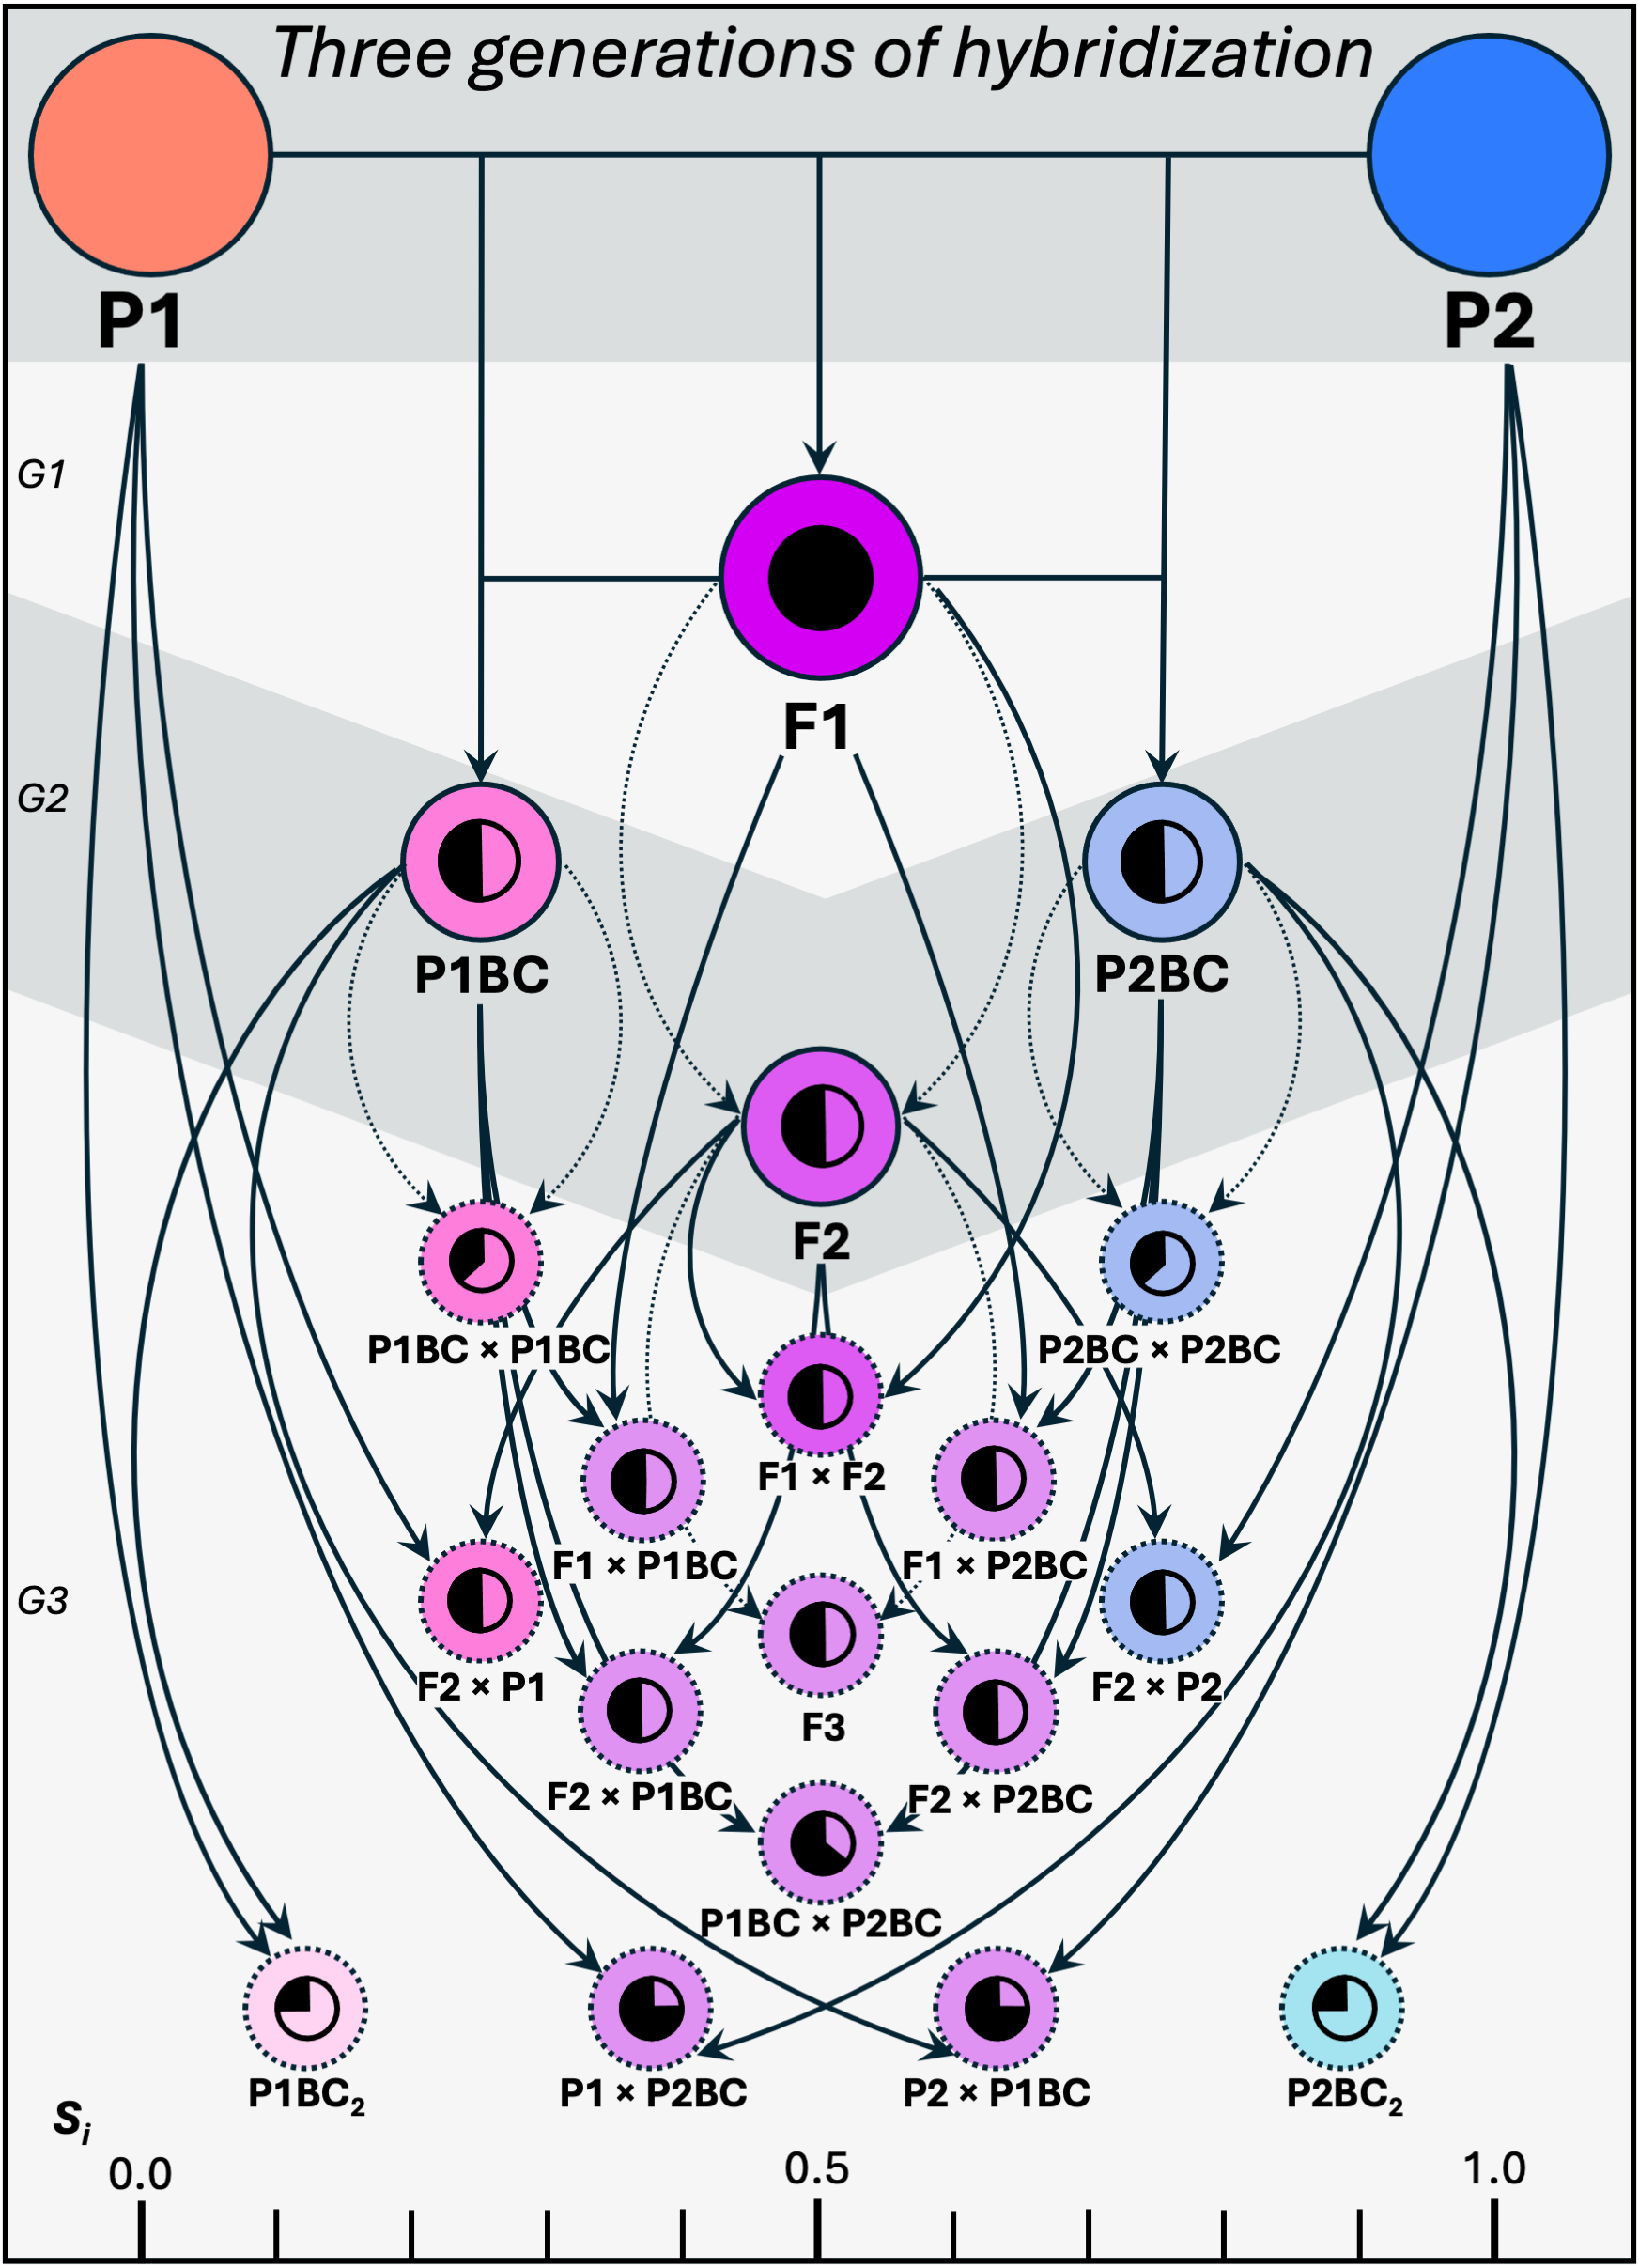
Hybridization is real and ongoing. Of the toads tested in the contact zone, 88% were backcrossed hybrids carrying an average of 6.5% Yosemite toad ancestry, with an estimated origin fewer than ten generations back. Read alongside the 1956 observation, that suggests new hybrids appear every five or six generations, followed by backcrossing to western toads every two to four. We also found trace Yosemite toad ancestry well outside any known contact zone, and trace western toad ancestry inside Yosemite and Kings Canyon National Parks, pointing to older introgression nobody had detected.
Whether this is a conservation problem is genuinely unclear, and the report says so. Novel variation entering a genetically depauperate species could fuel future adaptation; it could equally disrupt existing local adaptation. There is no evidence of genetic swamping or range replacement in the current data.
Technical details
225 new samples across four putative hybrid locations and ten western toad locations were merged with 644 existing Yosemite toad samples, for 877 individuals total. ddRADseq libraries reproduced the original digestion enzymes and fragment size window exactly, which was essential for locus compatibility across datasets. Sequencing ran on an Element AVITI.
Processing used STACKS v2.66, giving a final set of 3,814 loci across 866 individuals
after missingness filtering. Ancestry proportions came from ADMIXTURE at K=2, hybrid
classes from NEWHYBRIDS on 200 near-fixed markers, and joint estimates of hybrid index and
interspecies heterozygosity from HIest. Hybrid age was estimated by simulating 100
generations of a hybrid swarm and matching observed values to kernel density estimates per generation,
then taking a weighted mean of ages at the inflection of the ranked score curve.
Related work
The fungus
Chytridiomycosis, caused by Batrachochytrium dendrobatidis, has driven amphibian declines worldwide, including in this species. Pathogens in the field experience temperatures that swing widely over a single day, which constant-temperature lab cultures do not capture. Using temperature data logged from actual Yosemite toad breeding pools, we grew the fungus under a realistic daily fluctuation and found reduced growth, reduced zoospore production, and reduced zoosporangia viability compared with a constant regime at the same mean temperature.
A conifer in China
The landscape genomics methods transfer across taxa. Applied to Platycladus orientalis, a widespread Chinese conifer of ecological, timber, and medicinal value, genotyping-by-sequencing across the species range produced over 11,000 high-quality variants, evidence of isolation by environment, and estimates of genetic offset under future climate scenarios. Gradient forest modeling identified temperature variables as the strongest predictors, which bears directly on seed source selection for reforestation.
A desert vole
The Amargosa vole is an endangered rodent restricted to wetland vegetation along about 3.5 km of the Amargosa River in the Mojave Desert. We quantified its abundance, occupancy, and habitat selection across gradients of vegetation cover and spatial scale. This is the sort of baseline demographic work any recovery plan has to rest on, and it is a reminder that not every conservation question is a genomics question.
Natural history and ongoing work
Some observations are worth publishing on their own. Two short notes describe Yosemite toad larval diet and a breeding behavior observation, the latter with video. A collaboration on North American water mites, using phylogenomic methods on a group where morphology has been misleading, is in preparation, as is a paper on larval development evolution among pure and admixed lineages.
Field sites
Yosemite and Kings Canyon National Parks are my primary study areas, together comprising about 40% of the Yosemite toad distribution. Not a bad place for science. Roughly 99% of Yosemite toad breeding sites are meadows, either in mid-elevation montane forest or in subalpine rocky moonscape. Across eight field seasons I have spent more than 3,000 hours in them, most of it walking between one and the next. More photographs of the meadows are here.
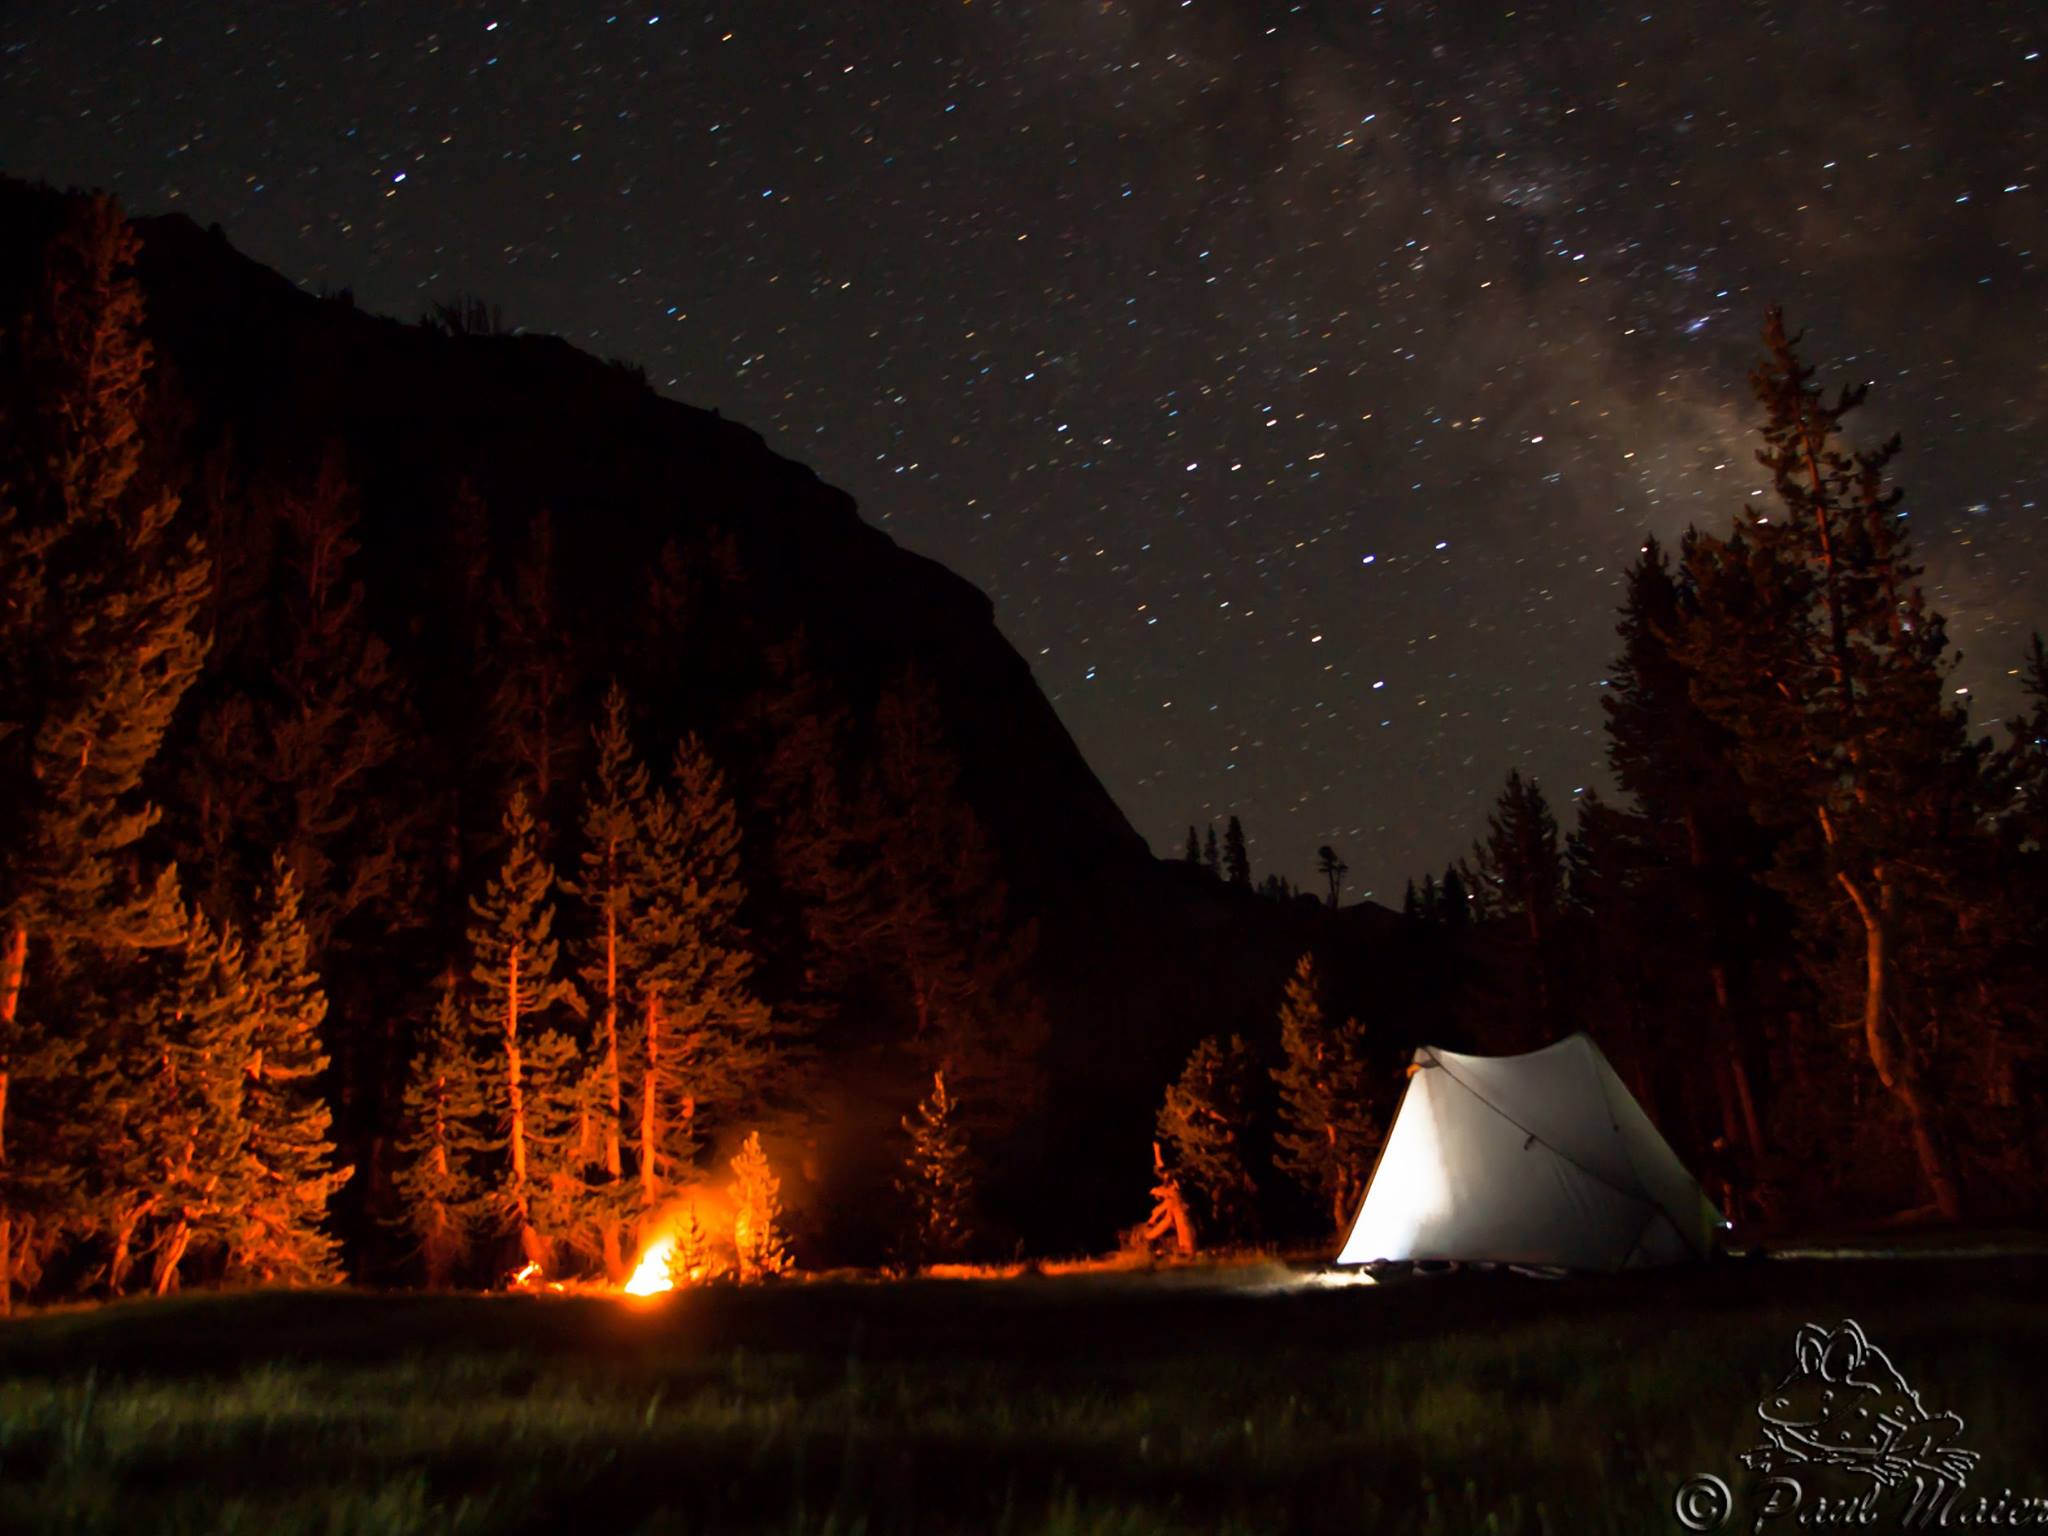
Below is the Milky Way from upper Kerrick Canyon in northern Yosemite in 2016, right before the peak of the Perseids. Northern Yosemite is remote and difficult to reach for the average recreational hiker. Starving graduate students and unpaid interns are not average.
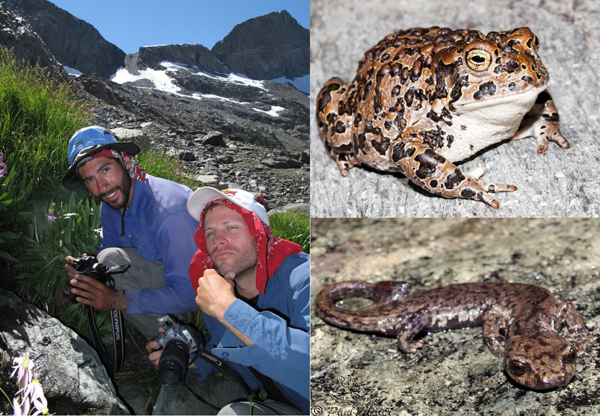
Kings Canyon holds the southern extent of the species and is my other primary site. Below is Goddard Canyon, where my field assistant Ross Maynard and I collected tadpole tail tips in 2011 (left). We took a break to photograph two rare finds: an adult female toad (top right) and a Mount Lyell salamander (Hydromantes platycephalus, bottom right).
Papers from this work
- Maier P.A., Vandergast A.G., Bohonak A.J. (2024). Yosemite toad transcriptome reveals interplay between speciation genes and adaptive introgression. Molecular Ecology 33(8), e17317. DOI · PDF
- Maier P.A., Mabe J.A., Hale J.L., Herman A.E., Kantz B.P. (2024). Hybridization between Anaxyrus boreas × canorus within Eldorado National Forest: the first genomic assessment. Final Report to the US Forest Service. DOI · Report
- Maier P.A., Vandergast A.G., Bohonak A.J. (2023). Using landscape genomics to delineate future adaptive potential for climate change in the Yosemite toad. Evolutionary Applications 16(1), 74–97. DOI · PDF
- Maier P.A., Vandergast A.G., Ostoja S.M., Aguilar A., Bohonak A.J. (2022). Landscape genetics of a sub-alpine toad: climate change predicted to induce upward range shifts via asymmetrical migration corridors. Heredity 129(5), 257–272. DOI · PDF
- Maier P.A., Vandergast A.G., Ostoja S.M., Aguilar A., Bohonak A.J. (2022). Gene pool boundaries for the Yosemite toad reveal asymmetrical migration within meadow neighborhoods. Frontiers in Conservation Science 3, 851676. DOI · PDF
- Maier P.A., Vandergast A.G., Ostoja S.M., Aguilar A., Bohonak A.J. (2019). Pleistocene glacial cycles drove lineage diversification and fusion in the Yosemite toad. Evolution 73(12), 2476–2496. DOI · PDF
- Lindauer A.L., Maier P.A., Voyles J. (2020). Daily fluctuating temperatures decrease growth and reproduction rate of a lethal amphibian fungal pathogen in culture. BMC Ecology 20(18). DOI · PDF
- Jia K.-H., Zhao W., Maier P.A., et al. (2020). Landscape genomics predicts climate change vulnerability for a forest tree. Evolutionary Applications 13(4), 665–676. DOI · PDF
- Klinger R., Cleaver M., Anderson S., Maier P., Clark J. (2015). Implications of scale-dependent habitat specialization on persistence of a rare small mammal. Global Ecology and Conservation 3, 100–114. DOI · PDF
- Lee S.R., Ostoja S.M., Maier P.A., et al. Distribution and spatiotemporal variation of Yosemite toad populations across Sierra Nevada national parks. Herpetological Conservation and Biology, in review.
- Maier P.A. (2018). Evolutionary past, present, and future of the Yosemite toad. Ph.D. dissertation, UC Riverside and San Diego State University. eScholarship